Overview of Validation Requirements 
in 
Pharmaceutical Industry 
Kaushik Desai 
Chairman, 
Industrial Pharmacy Division 
Indian Pharmaceutical Association 
www.ipapharma.org 
1
 A definition 
 Where did validation come from ? 
 Why do it ? 
 What are the benefits ? 
 How far do we have to go ? 
2
“ Establishing the documented evidence which 
provides a high degree of assurance that a 
specific process will consistently produce a 
product of predetermined specifications and 
quality attributes.” 
(FDA Guidelines 1987) 
3
“Process validation is defined as the collection 
& evaluation of data, from the process design 
stage through commercial production, which 
establishes scientific evidence that process is 
capable of consistently delivering quality 
product.” 
(FDA Guidelines, 2011) 
4
“Action of proving, in accordance with the 
principles of Good Manufacturing Practice 
(GMP), that any procedure, process, 
equipment, material, activity or system 
actually leads to expected results.” 
(EU GMP 1997) 
5
 Began in 1970’s 
 Originally sterilized based. 
 Now evolved into all Product, Process and Facility 
matters. 
6
 Assures Quality 
 Regulatory Requirement 
 Reduces Cost 
 It’s the LAW ! 
7
Validation can reduce costs by reducing, 
 Rejects 
 Reworks 
 Reliance on In-process controls 
 Down time 
8
 Reliance on product testing 
 Loss of confidence 
 Possibility of adulterated products 
 Inspection : Observations / 483’s 
9
 User Requirement Specification (URS) 
 Design Qualification 
 Impact Assessment 
 Factory Acceptance Testing 
 Installation Qualification 
10
 Calibration 
 Site Acceptance Testing 
 Operational Qualification 
 Standard Operating Procedures 
 Performance Qualification 
 Process Validation 
 Change Control 
11
User Requirement Specification (URS) 
 A description of the requirements of the facility 
(project) in terms of product to be manufactured, 
required throughput and conditions in which the 
product should be manufactured. 
 Approved statements prepared by the user which 
defines what is required by the project. 
12
Design Qualification (DQ) 
 Documented review of the design, at an 
appropriate stage in a project, for conformance 
to operational and regulatory expectations. 
(Note : Not an obligation) 
13
Impact Assessment 
 The process of evaluating the impact of the 
operating, controlling, alarming and failure 
conditions of a system on the quality of a 
product 
14
Factory Acceptance Testing (FAT) 
 Inspection and static and/or dynamic testing of 
systems or major system components to 
support the qualification of an equipment 
system conducted and documented at a 
supplier site. 
(Note : Not an obligation) 
15
 Documented verification that all aspects of a facility, 
utility or equipment that can affect the product 
quality adhere to approved specifications and are 
correctly installed. 
 The process of checking/verifying the installation to 
ensure that the critical components meet the 
approved specifications and that they are installed 
correctly in accordance with design documentation. 
16
 To establish that the critical components are installed 
correctly and in accordance with design 
documentation requirements (i.e. PO’s, Contracts 
etc.), that supporting documentation is in place and of 
suitable quality. 
 To record the checks and verifications for critical 
components in Direct Impact Systems. 
17
 Approval Page 
 Objectives 
 System Description 
 Responsibilities 
 Acceptance Criteria 
 Engineering Documentation Requirements 
 Records of Signatures 
 Qualification Test Equipment/Instrument List 
 Product Contact Materials Review 
 Utilities Verification 
 Control System Verification 
18
 Instrument/Control Devices Verification 
 Equipment Verification 
 Piping Installation Verification 
 Discrepancy/Justification and corrective Action 
 As built P&I Diagrams 
 Specifications 
 Conclusions 
 References 
 Modification/ Change Control 
 Attachments / Appendices 
19
Parameters 
 Does the vessel meet the design specification? 
 Does the agitator assembly meet the design specification? 
 Is the motor housing earthed? 
 Is the motor over current device set to correct setting? 
 Is all the pipe work connected? 
 Are all instruments installed as per P&I diagram? 
 Have all the temperature indicators been calibrated? 
 Is calibration procedure available? 
 Are operation and maintenance manuals available? 
 all electrical connections securely and safely fitted? 
 Is insulation complete? 
 Is vessel clean and free from dirt? 
20
 After protocol execution is complete and 
deviations evaluated, post execution approval 
is required. 
 Requires sign off by original signatories. 
 IQ execution should be complete and approved 
prior to the start of OQ. 
21
Calibration 
 Demonstrating that a measuring device 
produces results within the specified limits of 
those produced by a reference standard over 
an appropriate range of measurements. 
 The devices are normally tagged and 
supported by a maintenance procedures. 
22
Site Acceptance Testing (SAT) 
 Inspection and dynamic testing of systems or 
major system components to support the 
qualification of an equipment system 
conducted at a client site. 
(Note : Not an obligation) 
23
 Documented verification that all aspects of 
a facility, utility or equipment that can affect 
product quality operate as intended 
throughout all anticipated ranges. It is the 
process of testing to ensure that individual 
components and systems operate as 
specified, and how that information is 
recorded. 
24
 To establish through documented testing, that all 
critical components and direct impact systems are 
capable of operating within established limits and 
tolerances. 
 To test parameters that regulate the process or 
product quality. To verify the proper operation of 
controllers, indicators, recorders, alarms and 
interlocks, is performed and documented during the 
operational qualification testing. 
25
 Approval page 
 Pre-requisites 
 Objectives 
 System Description 
 Responsibilities 
 Acceptance Criteria 
 Records of signatures 
 Qualification test Equipment/Instruments list 
 Alarm and Interlocks test 
 Operation testing 
 Capacity testing 
 Power failure testing 
26
 Sequence testing 
 Test data sheets 
 SOP’s 
 Conclusions 
 Modification / change control 
 Discrepancy/Justification and corrective action 
 Operational Qualification Summary 
 References 
 Attachments/Appendices 
- Verification of test instruments 
- Chart recordings 
- P&I diagrams 
- Printouts 
27
Parameters 
 Have all Installation Qualification been completed for this system? 
 Is the system clean and free from dirt? 
 Is the direction of the rotation of agitator correct? 
 Check the operation of the agitator emergency stop? 
 Check the operation of all agitator controls, both on the main and local 
panels? 
 Check that the agitator in the vessel free to turn? 
 Pressurize the vessel and record the pressure drop for 10 min. 
 Perform a vacuum test and record the vacuum drop. 
28
 Documented verification that all aspects of 
a facility, utility or equipment that can 
affect the product quality perform as 
intended in meeting the predetermined 
acceptance criteria. 
29
 To integrate procedures, personnel, systems 
and materials to verify that the utility / 
environment / equipment / support systems 
produces the required output. This output 
may be a product contact utility, sterilization 
condition or environment. 
30
 Approval page 
 Pre-requisites 
 Objectives 
 System Description 
 Responsibilities 
 Acceptance Criteria 
 PQ test plan 
 Challenge test plan 
 Records of signatures 
 Test equipment/Instrument list 
 Test data sheets 
 SOP’s 
 References 
 Conclusions 
 Attachments 
31
Process Validation 
 The documented verification providing a high 
degree of assurance that a specific process will 
consistently produce a product meeting its 
predetermined specifications and quality 
attributes. 
 The new guidelines aligns process validation 
activities with a product life cycle concept. 
32
 211.42, 211.63, 211.68, 211.84 
 211.100(a) , 211.110(a), 
211.110(b) 
 211.160(b), 211.165(c), 
211.165(d), 211.180(e) 
33
Life Cycle of the product & the process 
 Stage 1 – Process Design 
 Stage 2 – Process qualification 
 Stage 3 - Continuous process verification 
34
 It is the activity of defining the commercial manufacturing 
process that will be reflected in planned master production 
and control records. 
 The goal of this stage is to design a process suitable for 
routine commercial manufacturing that can consistently 
deliver a product that meets its quality attributes. 
 It is based on the knowledge gained through development & 
scale-up activities. 
35
During this stage, the process design is evaluated to 
determine if the process is capable of reproducible 
commercial manufacturing. The products 
manufactured during this stage, if acceptable , can be 
released for distribution. 
Two Aspects 
 Design of facility and qualification of equipment and 
utilities. 
 Process Performance Qualification ( PPQ). 
36
Def.: “Releasing for distribution a lot of finished 
product, manufactured following a qualification 
protocol, that meets the standards established in 
the protocol, but before the entire study has been 
executed”. 
 Orphan Drugs 
 Specific drug for specific use 
 Shor t Shelf-life radio pharmaceuticals 
37
 The goal of this stage is continual assurance that the 
process remains in a state of control ( the validated state) 
during commercial manufacture. 
 The cGMP requirements, specifically the collection & 
evaluation of information & data about the performance 
of the process will allow detection of undesired process 
variability. 
 This stage is also applicable for legacy products. 
38
Change Control 
 Formal evaluation of the potential impact of 
planned modifications on the validated status 
of a product, process or facility. 
39
 Training personnel for IQ/OQ execution. 
 The purpose of the equipment/ system. 
 Use of test equipment 
 Applicable SOP’s 
 cGMP documentation of training 
 Document all training 
 Periodically review training requirements 
40
 A document that summarizes and analyses 
the test results at the end of PQ. 
 Provides a conclusion about the ability of the 
system to consistently meet acceptance 
criteria. 
 May be a stand alone document at each 
stage of IQ/OQ and PQ to summarize results. 
41
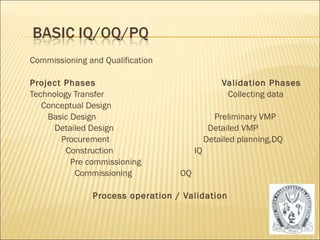
Commissioning and Qualification 
Project Phases Validation Phases 
Technology Transfer Collecting data 
Conceptual Design 
Basic Design Preliminary VMP 
Detailed Design Detailed VMP 
Procurement Detailed planning,DQ 
Construction IQ 
Pre commissioning 
Commissioning OQ 
Process operation / Validation 
42
43 
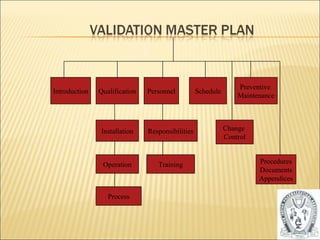
Introduction Qualification Personnel Schedule Preventive 
Maintenance 
Installation 
Operation 
Process 
Responsibilities 
Training 
Change 
Control 
Procedures 
Documents 
Appendices
The company’s overall policy, intentions and 
approach to validation, including : 
 Validation of production processes 
 Cleaning procedures 
 Analytical methods 
 In-process control test procedures 
 Computerized systems 
 Persons responsible for design, review, approval 
 Documentation of each validation phase 
44
 Product validation is NOT …… 
- just 3 batches that meet specifications 
 Product validation is …… 
- an ongoing process to build confidence into 
the manufacturing activities 
- an ability to demonstrate consistency at any 
time. 
45
 Cleaning validation is establishing 
documented evidence that the equipment 
is consistently cleaned from product, 
microbial and cleaning agent residues to 
predetermined acceptable levels. 
46
A written procedure for cleaning validation 
which includes : 
 Responsibility for development, performance and 
approval of the validation study. 
 Establishment of SOP’s 
 Acceptance criteria 
- defined to prevent cross contamination 
- definition of residue limits 
47
 Cleaning procedure for each piece of equipment 
- Flow charts and diagrams 
- Cleaning agents, concentration, volume 
- Frequency 
- time left ‘dirty’ 
48
 Sampling procedures 
- swabs, rinse, location 
 Residue limits 
 Analytical methods 
49
 Validation report 
- Results Vs. acceptance criteria 
- Deviations and how handled 
- Conclusion that cleaning process is 
validated 
50
 Major change in cleaning procedure 
 Change in cleaning agent 
 New equipment 
51
 Knowledge of the materials 
- Potency of the drug 
- Pharmacological and toxic properties 
- Degradation products 
- Cleaning agents 
- Micro residues 
52
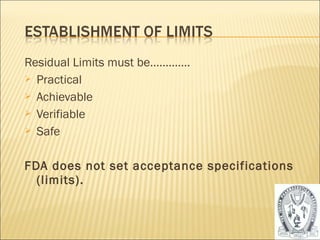
Residual Limits must be…………. 
 Practical 
 Achievable 
 Verifiable 
 Safe 
FDA does not set acceptance specifications 
(limits). 
53
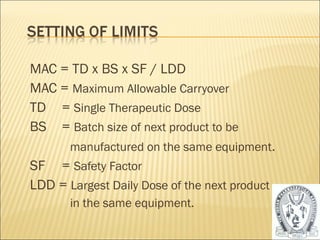
MAC = TD x BS x SF / LDD 
MAC = Maximum Allowable Carryover 
TD = Single Therapeutic Dose 
BS = Batch size of next product to be 
manufactured on the same equipment. 
SF = Safety Factor 
LDD = Largest Daily Dose of the next product 
in the same equipment. 
54
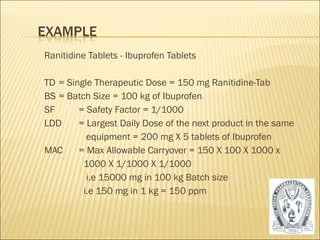
Ranitidine Tablets - Ibuprofen Tablets 
TD = Single Therapeutic Dose = 150 mg Ranitidine-Tab 
BS = Batch Size = 100 kg of Ibuprofen 
SF = Safety Factor = 1/1000 
LDD = Largest Daily Dose of the next product in the same 
equipment = 200 mg X 5 tablets of Ibuprofen 
MAC = Max Allowable Carryover = 150 X 100 X 1000 x 
1000 X 1/1000 X 1/1000 
i.e 15000 mg in 100 kg Batch size 
i.e 150 mg in 1 kg = 150 ppm 
55
Validation