2 
Objectives: 
 Dissolution science is not just a quality control tool. 
Apart from that in present era dissolution data act as 
surrogate marker for in-vivo bioavailability . 
 Along with wide versatility of application to 
pharmaceutical scientist it also form basis for setting 
specification to allow the release of batch to market. 
 Present seminar try’s to give a bird eye view of 
various dissolution model’s which will be helpful in 
predicting the drug release kinetics ( dissolution 
kinetics.)
DISSOLUTION 
Dissolution is a process in which a solid substance 
solubilizes in a given solvent i.e. mass transfer from 
the solid surface to the liquid phase 
Rate of dissolution is the amount of drug substance 
that goes in solution per unit time under standardized 
conditions of liquid/solid interface, temperature and 
solvent composition 
3
Why dissolution studies? 
1. To show that the release of drug from the 
tablet is close to 100%. 
2. To show that the rate of drug release is 
uniform batch to batch. 
3. And to show that release is equivalent to 
those batches proven to be bioavailable 
and clinically effective. 
4
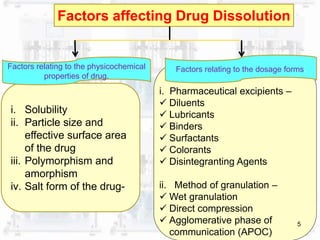
Factors affecting Drug Dissolution 
Factors relating to the physicochemical 
properties of drug. 
i. Solubility 
ii. Particle size and 
effective surface area 
of the drug 
iii. Polymorphism and 
amorphism 
iv. Salt form of the drug- 
Factors relating to the dosage forms. 
i. Pharmaceutical excipients – 
 Diluents 
 Lubricants 
 Binders 
 Surfactants 
 Colorants 
 Disintegranting Agents 
ii. Method of granulation – 
 Wet granulation 
 Direct compression 
 Agglomerative phase of 
communication (APOC) 
5
DISSOLUTION MODEL’s & IT’s NEED 
 Dissolution Profile- It is graphical representation [in 
terms of concentration vs. time] of complete release of 
A.P.I. from a dosage form in an appropriate selected 
dissolution medium. 
i.e. in short it is the measure of the release of A.P.I 
from a dosage form with respect to time. 
 IT’s NEED 
 To Develop invitro-invivo correlation which can 
help to reduced costs, speed-up product 
development and reduced the need of perform costly 
bioavailability human volunteer studies. 
 To stabilize final dissolution specification for 
the pharmacological dosage form 
6
TYPES OF DISSOLUTION MODELS 
1 • Diffusion layer model 
2 • Danckwert’s MODEL 
3 • Interfacial barrier model 
4 • Zero-order model 
5 • First-order model 
6 • Higuchi model 
7 • Korsmeyer-Peppas model 
8 • Hixson-Crowell model 
9 • Baker-Lonsdale model 
10 • Weibull model 7
1. DIFFUSION LAYER MODEL 
 Formation of a thin film at the interface, called as stagnant 
layer. 
 2 steps are involved: 
i. Interaction of solvent with drug surface to form a saturated 
drug layer , called stagnant layer. 
ii. Diffusion of drug molecules from stagnant layer into bulk of 
the system.
1. DIFFUSION LAYER MODEL (contd…) 
Using Fick’s law, Noyes-Whitney equation for DIFFUSION LAYER 
MODEL is as follows 
Where, 
 dC/dt = dissolution rate of the drug. 
 D = diffusion coefficient of the drug. 
 A = surface area of the dissolving solid 
 Kw/o = water/oil partition coefficient of drug 
 V = volume of dissolution medium 
 h = thickness of stagnant layer 
 Cs–Cb = concentration gradient of diffusion of drug 
9
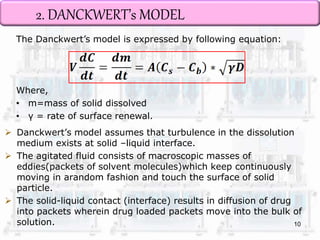
2. DANCKWERT’s MODEL 
The Danckwert’s model is expressed by following equation: 
Where, 
• m=mass of solid dissolved 
• ү = rate of surface renewal. 
 Danckwert’s model assumes that turbulence in the dissolution 
medium exists at solid –liquid interface. 
 The agitated fluid consists of macroscopic masses of 
eddies(packets of solvent molecules)which keep continuously 
moving in arandom fashion and touch the surface of solid 
particle. 
 The solid-liquid contact (interface) results in diffusion of drug 
into packets wherein drug loaded packets move into the bulk of 
solution. 
10
2. DANCKWERT’s MODEL (contd…) 
 In agitated fluid drug loaded packets are continuously replaced by 
fresh packets due to which drug conc at solid liquid interface 
never reaches saturation concentration. 
 Since the solvent packets continuously replace the surface of the 
solid , it is called as SATURATION RENEWAL THEORY 
11
3. INTERFACIAL BARRIER MODEL 
 Interfacial barrier model considers drug dissolution as crystal 
dissolution wherein solids get hydrated initially and is not 
instantaneous 
 The reaction at solid surface and its diffusion across the interface 
is slower than diffusion across liquid film 
 Therefore the rate of solubility of solid in liquid film becomes the 
rate limiting than the diffusion of dissolved molecules 
 When considering the dissolution of the crystal will have a different 
interfacial barrier given by following equation, 
G = ki (Cs – Cb) 
 Where, 
 G = dissolution per unit area 
 Ki = effective interfacial transport constant 
12
4. Zero-order model 
 Drug dissolution from dosage forms that do not disaggregate and 
release the drug slowly can be represented by the equation: 
Q0-Qt = K0t 
Rearrangement of equation yields: 
Qt = Q0 + K0t 
where , 
 Qt is the amount of drug dissolved in time t, 
 Q0 is the initial amount of drug in the solution (most times, Q0 = 
0) and K0 is the zero order release constant expressed in units of 
concentration/time. 
The release rate 
is independent 
of concentration 
13
5. First-order model 
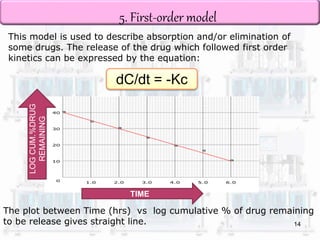
This model is used to describe absorption and/or elimination of 
some drugs. The release of the drug which followed first order 
kinetics can be expressed by the equation: 
LOG CUM.%DRUG 
REMAINING 
dC/dt = -Kc 
TIME 
The plot between Time (hrs) vs log cumulative % of drug remaining 
to be release gives straight line. 
14
6. HIGUCHI MODEL 
 Higuchi developed models to study the release of water 
soluble and low soluble drugs incorporated in semisolid 
and solid matrices. 
 To study the dissolution from a planar system having a 
homogeneous matrix the relation obtained was; 
A = [D (2C – Cs)Cs × t]1/2 
Where, 
 A is the amount of drug released in time ‘t’ per unit 
area, 
 D is the diffusivity of drug molecules in the matrix 
substance 
 C is the initial drug concentration, 
 Cs is the drug solubility in the matrix media 
15
6. HIGUCHI MODEL (contd…) 
Applications: 
 Higuchi describes the drug release as a diffusion process based 
on Ficks law, square root time dependent . 
 This model is useful for studying the release of water soluble and 
poorly soluble drugs from variety of matrices ,including solids 
and semi solids. 
16
7. Hixson-Crowell model 
 Hixon-Crowell recognized that the particle regular area is 
proportional to the cubic root of its volume, and hence desired 
an equation as 
1/3- M1/3 = K × t 
Mo 
where, Mo = original mass of particles 
K = cube-root dissolution rate constant 
M = mass of the A.P.I at the time ‘t’ 
APPLICATIONS: 
 To evaluate the drug release with changes in the surface area 
and the diameter of the particles /tablets 
 The rate of dissolution depends on the surface of solvent - the 
larger is area the faster is dissolution. 
 It describes the drug releases by dissolution, with the changes 
in surface area and diameter of the particles or tablets. 
17
7. Hixson-Crowell model (contd…) 
 The plotted graph will be linear if the following conditions 
are fulfilled:- 
 The equilibrium conditions are not reached and 
 The geometrical shape of the pharmaceutical dosage form 
diminishes proportionally over time. 
18
8. Korsmeyer-Peppas model 
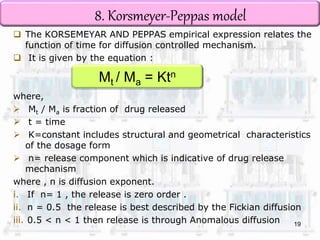
 The KORSEMEYAR AND PEPPAS empirical expression relates the 
function of time for diffusion controlled mechanism. 
 It is given by the equation : 
Mt / Ma = Ktn 
where, 
 Mt / Ma is fraction of drug released 
 t = time 
 K=constant includes structural and geometrical characteristics 
of the dosage form 
 n= release component which is indicative of drug release 
mechanism 
where , n is diffusion exponent. 
i. If n= 1 , the release is zero order . 
ii. n = 0.5 the release is best described by the Fickian diffusion 
iii. 0.5 < n < 1 then release is through Anomalous diffusion 
19
10.Korsmeyer-Peppas model (contd…) 
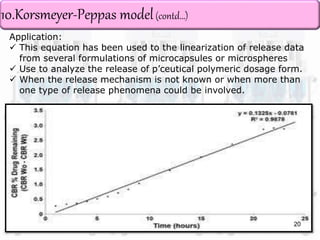
Application: 
 This equation has been used to the linearization of release data 
from several formulations of microcapsules or microspheres 
 Use to analyze the release of p’ceutical polymeric dosage form. 
 When the release mechanism is not known or when more than 
one type of release phenomena could be involved. 
20
9.Baker-Lonsdale model 
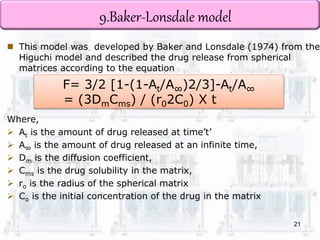
 This model was developed by Baker and Lonsdale (1974) from the 
Higuchi model and described the drug release from spherical 
matrices according to the equation 
F= 3/2 [1-(1-At/A∞)2/3]-At/A∞ 
= (3DmCms) / (r02C0) X t 
Where, 
 At is the amount of drug released at time’t’ 
 A∞ is the amount of drug released at an infinite time, 
 Dm is the diffusion coefficient, 
 Cms is the drug solubility in the matrix, 
 ro is the radius of the spherical matrix 
 Co is the initial concentration of the drug in the matrix 
21
10. Weibull model 
Weibull model is generally applied to drug dissolution or 
release from pharmaceutical dosage forms. The accumulated 
fraction of the drug M in solution at time t is given by Weibull 
equation: 
M = M0[1-e-(t-T/a)b] 
Where, 
 m = % dissolved at time ‘t’ 
 a = scale parameter which defines time scale of the 
dissolution process 
 T1 = location parameters which represents lag period 
before the actual onset of dissolution process (in most of 
the cases T1 = 0) 
 b = shape parameter which quantitatively defines the curve 
Application: 
The Weibull model is more useful for comparing the release 
profiles of matrix type drug delivery 
22
1) Remington's “The science and practice of pharmacy” 21st 
edition page no 672-685. 
2) “A Text book of Applied Bio pharmaceutics and 
pharmacokinetics”, by Leon Shargel,andrew , 4 th edition 
,page no 131-195. 
3) “Text book of Bio pharmaceutics and pharmacokinetics” 
,by V.Venkateshwarlu page no.32-55. 
4) “Text book of Bio pharmaceutics and pharmacokinetics”, 
by Brahmankar.page no.15-48. 
5) European Journal of Pharmaceutical sciences 13 (2001) 
page no.123 – 133. 
6) Mathematical models of dissolution- Master’s thesis by 
Jakub ˘ Cupera May 4, 2009 Masarykova Univerzita 
7) Vinod P. Shah The role of dissolution testing in the 
regulation of pharmaceuticals: The FDA perspective, 
Taylor and Francis Group 2005 page no.81-95. 
23 
References
CONCLUSION 
 The Quantitative interpretation of the values 
obtained n dissolution assays is easier using 
mathematical equations which describe the release 
profile in function of some parameters related with 
the pharmaceutical dosage forms. 
 The release models with the major appliance and 
the best describing drug release phenomena. 
 The Higuchi model has a larger application in 
polymeric systems, the zero order model becomes 
ideal to describe coated dosage forms or membrane 
controlled dosage forms. 
24
25