By D. Saidulu
Rule

Permission for

122 A

To import New drugs

122 B

To manufacture New drugs

122 D

To Import or Manufacture fixed
dose combinations

122DA

To conduct clinical trials for New
Drug/Investigational New drug

122 DAA

Definition of Clinical Trial
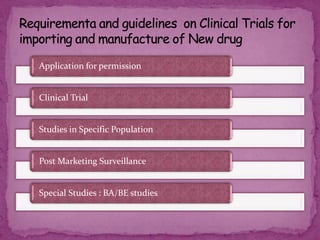
Application for permission

Clinical Trial

Studies in Specific Population

Post Marketing Surveillance

Special Studies : BA/BE studies
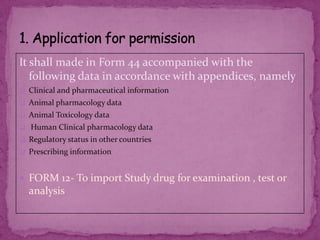
It shall made in Form 44 accompanied with the
following data in accordance with appendices, namely
 Clinical and pharmaceutical information
 Animal pharmacology data
 Animal Toxicology data
 Human Clinical pharmacology data

 Regulatory status in other countries
 Prescribing information

 FORM 12- To import Study drug for examination , test or

analysis
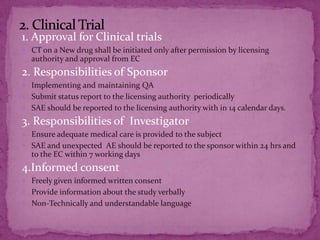
1. Approval for Clinical trials
 CT on a New drug shall be initiated only after permission by licensing

authority and approval from EC

2. Responsibilities of Sponsor
 Implementing and maintaining QA
 Submit status report to the licensing authority periodically
 SAE should be reported to the licensing authority with in 14 calendar days.

3. Responsibilities of Investigator
 Ensure adequate medical care is provided to the subject
 SAE and unexpected AE should be reported to the sponsor within 24 hrs and

to the EC within 7 working days

4.Informed consent
 Freely given informed written consent

 Provide information about the study verbally
 Non-Technically and understandable language
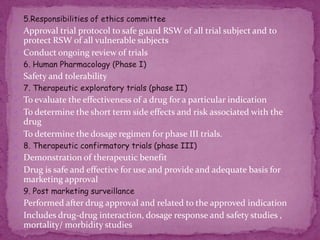
 5.Responsibilities of ethics committee

 Approval trial protocol to safe guard RSW of all trial subject and to

protect RSW of all vulnerable subjects
 Conduct ongoing review of trials
 6. Human Pharmacology (Phase I)

 Safety and tolerability
 7. Therapeutic exploratory trials (phase II)

 To evaluate the effectiveness of a drug for a particular indication
 To determine the short term side effects and risk associated with the

drug
 To determine the dosage regimen for phase III trials.
 8. Therapeutic confirmatory trials (phase III)

 Demonstration of therapeutic benefit
 Drug is safe and effective for use and provide and adequate basis for

marketing approval
 9. Post marketing surveillance

 Performed after drug approval and related to the approved indication
 Includes drug-drug interaction, dosage response and safety studies ,

mortality/ morbidity studies
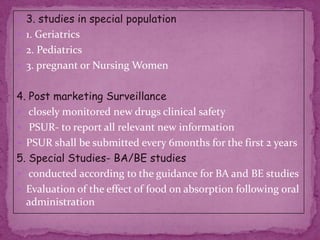
 3. studies in special population

 1. Geriatrics
 2. Pediatrics
 3. pregnant or Nursing Women

4. Post marketing Surveillance
 closely monitored new drugs clinical safety
 PSUR- to report all relevant new information
 PSUR shall be submitted every 6months for the first 2 years
5. Special Studies- BA/BE studies
 conducted according to the guidance for BA and BE studies
 Evaluation of the effect of food on absorption following oral
administration


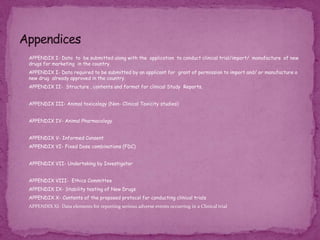
APPENDIX I- Data to be submitted along with the application to conduct clinical trial/import/ manufacture of new
drugs for marketing in the country.



APPENDIX I- Data required to be submitted by an applicant for grant of permission to import and/ or manufacture a
new drug already approved in the country.



APPENDIX II- Structure , contents and format for clinical Study Reports.



APPENDIX III- Animal toxicology (Non- Clinical Toxicity studies)



APPENDIX IV- Animal Pharmacology



APPENDIX V- Informed Consent



APPENDIX VI- Fixed Dose combinations (FDC)



APPENDIX VII- Undertaking by Investigator



APPENDIX VIII- Ethics Committee



APPENDIX IX- Stability testing of New Drugs



APPENDIX X- Contents of the proposed protocol for conducting clinical trials



APPENDIX XI- Data elements for reporting serious adverse events occurring in a Clinical trial
Schedule y
Schedule y