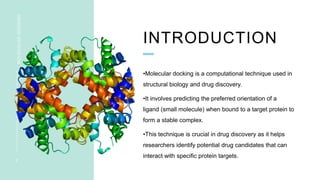
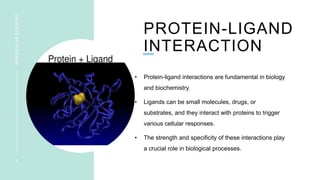
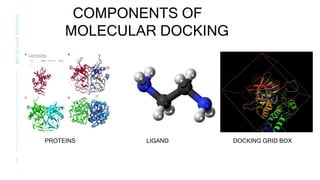
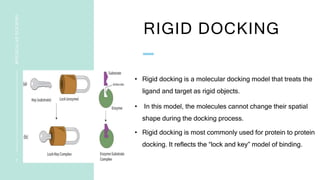
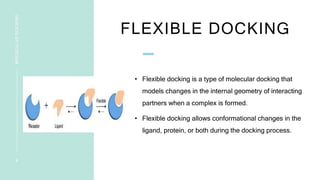
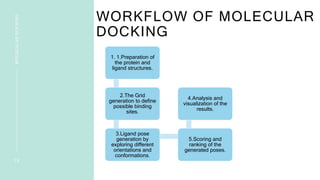
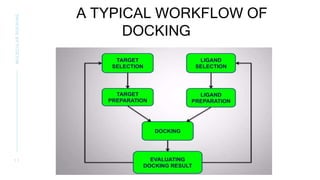
Molecular docking is a computational technique in structural biology and drug discovery that predicts the orientation of ligands binding to target proteins. It involves components such as protein and ligand structures, type of docking (rigid or flexible), and various interaction types, playing a crucial role in applications like drug design, virtual screening, and understanding protein-ligand interactions. Software tools like AutoDock and Glide are commonly used for these processes, facilitating the identification and optimization of potential drug candidates.