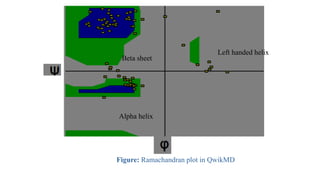
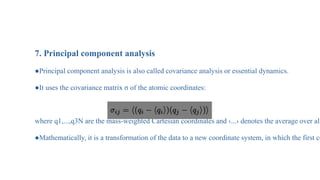
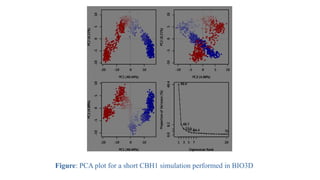
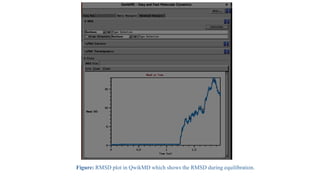
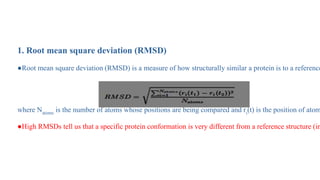
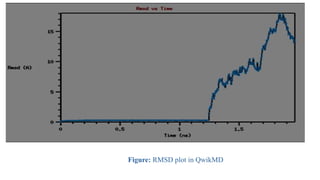
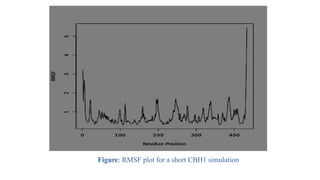
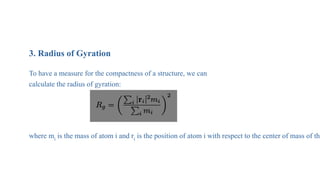
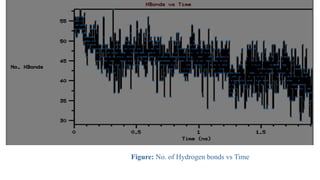
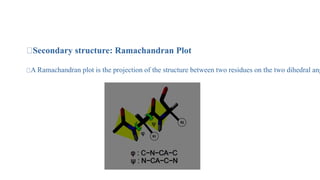
This document provides an overview of molecular dynamics (MD) simulations and their analysis. MD simulations calculate the time-dependent behavior of molecules and can be used to study conformational changes in proteins and nucleic acids. The document outlines various analyses that can be done on MD simulations including root mean square deviation (RMSD), root mean square fluctuation (RMSF), radius of gyration, hydrogen bonding, secondary structure analysis using Ramachandran plots, free energy surfaces, and principal component analysis. It also provides examples of running MD simulations using VMD and applications of MD simulations such as understanding allostery and molecular docking.


 Most read



 Most read