INVESTIGATION OF MEDICINAL
PRODUCT DOSSIER
Diptee Gupta
M.Pharm 1ST year
Pharmaceutics
CSJM University Kanpur 1
DRUG APPROVAL PROCESS-
There are 2 regulatory steps to go through before a drug is approved to be marketed in European Union
(EU).
The 2 steps are-
 Clinical trial application
 Marketing authorization application
 Clinical trial application are approved at the member state level where as Marketing authorization
application are approved both at member state or centralized level.
 There are some qualified persons who certify that INVESTIGATION OF MEDICINAL
PRODUCT DOSSIER (IMPD) should be manufactured according to GMP.
2
So definition of IMPD as –
“It is the basis for approval of clinical trials by the competent authority in the EU. The
clinical trial directive came in force to harmonizing the laws, regulations and administrative
provisions of member states relating to the implementation of GCP in the conduct of clinical
trials on medicinal products for human use.”
The directive introduce a harmonized procedure for authorization to perform clinical study in
any one of the EU member state. In addition it defines the investigation to be submitted to the
ethics committee as well as IMPD should be submitted to the competent authority for
approval.
3
DOSSIER
It defined as- “A collection of documents should a particular person, event or subject.”
eg. Patient Medical Dossier, M.Pharm Academic Dossier.
TYPES OF IMPD-
The EU has provided for two types of IMPDs, a “Full IMPD” and a “Simplified IMPD”, based
on whether the product has been described previously in another CTA or a marketing
authorization application. Guidance on the structure and content of an IMPD is provided by the
European Commission (EC) in ENTR/F2/BL D(2003) CT1 Revision 2, dated October 2005.
4
When applying for a clinical trial authorization-
 A full IMPD is required when little or no information about an IMP has been previously
submitted to competent authorities, when it is not possible to cross- refer to data submitted by
another sponsor and/or when there is no MA in the Community.
 A simplified IMPD may be submitted if information has been assessed previously as part of a
Marketing Authorization in any MS or a clinical trial to that competent authority.
Full and Simplified IMPD-
5
IMPDs are submitted as a part of Clinical trial application DOSSIER by competent
regulatory authorities within EU. It contain following parts-
 Protocols
 Informed consent forms
 Investigator brochure
 Study reports
 Subject narratives
 Risk management plan
Content of IMPD-
6
The IMPD required will depend on many factors including-
1. Risk aspects
2. Nature of product
3. State of development
4. Patient population
5. Nature and severity of illness
6. Type and duration of clinical trial itself
Thus IMPD also defined as-
“It is one of several pieces of investigational medicinal product (IMP) related data required
whenever the performance of clinical trial is intended in one or more EU member states.”
7
INVESTIGATOR BROUCHRE (IB)
Defined as-
“It is a compilation of clinical and non-clinical data on the investigational product that are
relevant to the study of products in human subjects.”
Purpose of IB-
 To provide information to the investigator and others involved in the trials such as dose,
dose frequency, dose interval, methods of administration and safety monitoring
procedures.
 Provide insight to support the clinical management for carryout clinical trial study.
 The information should be presented in a concise and simple manner.
 IB enable a clinician, or potential investigator, to understand it and make his own risk-
benefit assessment of the proposed trial. 8
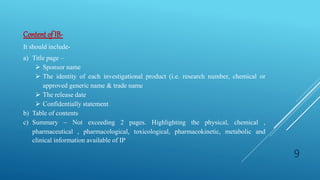
Contentof IB-
It should include-
a) Title page –
 Sponsor name
 The identity of each investigational product (i.e. research number, chemical or
approved generic name & trade name
 The release date
 Confidentially statement
b) Table of contents
c) Summary – Not exceeding 2 pages. Highlighting the physical, chemical ,
pharmaceutical , pharmacological, toxicological, pharmacokinetic, metabolic and
clinical information available of IP
9
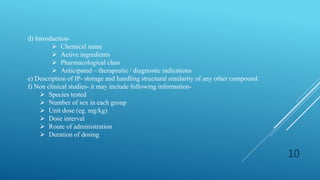
d) Introduction-
 Chemical name
 Active ingredients
 Pharmacological class
 Anticipated – therapeutic / diagnostic indications
e) Description of IP- storage and handling structural similarity of any other compound.
f) Non clinical studies- it may include following information-
 Species tested
 Number of sex in each group
 Unit dose (eg. mg/kg)
 Dose interval
 Route of administration
 Duration of dosing
10
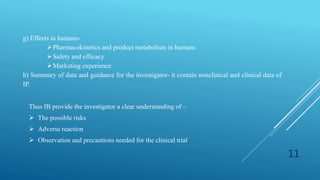
g) Effects in humans-
Pharmacokinetics and product metabolism in humans
Safety and efficacy
Marketing experience
h) Summary of data and guidance for the investigator- it contain nonclinical and clinical data of
IP.
Thus IB provide the investigator a clear understanding of –
 The possible risks
 Adverse reaction
 Observation and precautions needed for the clinical trial
11
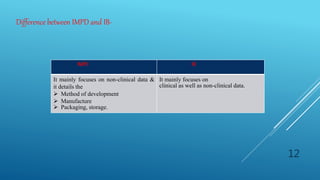
Difference between IMPD and IB-
IMPD IB
It mainly focuses on non-clinical data &
it details the
 Method of development
 Manufacture
 Packaging, storage.
It mainly focuses on
clinical as well as non-clinical data.
12
THANK-YOU
13