HATCH WAXMAN ACT
Learn Regulatory Affairs from Pharma IQ Education
Pharma IQ Education by Priyansha Singh (B. Pharm, M.S. Pharmacology & Toxicology)
Regulatory Affairs Executive- Viatris
CONTENTS
• Introduction
• Objectives of the act
• New Drug Approval
• Generic Drug Approval
• Challenging patent exclusivities
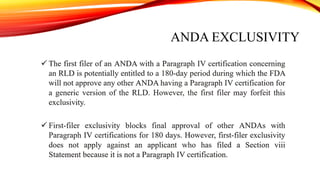
• ANDA exclusivity
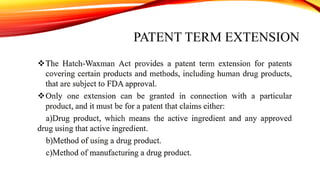
• Patent term extension
• Benefits for branded drug manufacturers
• Loopholes of the act
• Conclusion
OVERVIEW
• The "Drug Price Competition and Patent Term Restoration Act of 1984," also known as the Hatch-
Waxman Amendments, established the approval pathway for generic drug products, under which
applicants can submit an abbreviated new drug application (ANDA) under section 505(j) of the
Federal Food, Drug, and Cosmetic Act (FD&C Act).
• The Hatch-Waxman Amendments include provisions that involve patents and exclusivities related to
new drug applications, and 180-day exclusivity for certain ANDA applicants.
• As a general matter, the Food and Drug Administration (FDA or the Agency) has implemented these
statutory provisions within the context of application-specific decisions.
INTRODUCTION
• It is a.k.a. “The drug price competition & patent term restoration act”.
• Because the approval process for each new product required the submission of full safety and efficacy data, the
economic pressures resulting from the lengthy review process led to the Drug Price Competition and Patent
Restoration Act of 1984, also known as the Hatch-Waxman Act.
• This law established a process for the approval of drugs based on comparison to an already approved product
and provided for exclusive marketing status for a period of time based on the length of the approval process for
new drugs or the patent status of the branded drug for generics.
• Before 1962, approvals were done based alone on safety, but after 1962, proof of efficacy also was compulsory.
• Under the HWA act, a company can seek approval from the FDA to market a generic drug
before the patent expiry of the brand drug out of which the generic drug is based. The
company making generic drug files for an ANDA which after approval has the exclusive
right to market the generic drug for upto 180 days.
• There was no provision for the patent term extension before the enactment of HWA to cover
up for the time & money lost out of the total patent term during the marketing approval
process.
• Generic companies had to submit their own NDA which is costly & time consuming. Hence,
the act was required to solve these issues.
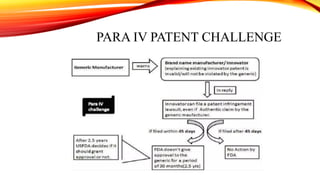
• To begin the FDA approval process, the generic applicant must
1. Certify in its ANDA that the patent in question is invalid/ is not infringed by the generic
product (a.k.a Paragraph IV certification); and
2. Notify the patent holder of the submission of the ANDA. If the patent holder files an
infringement suit against the generic applicant within 45 days of the ANDA notification, FDA
approval to market the generic is automatically postponed for 30 months, unless, before that
time the patent expires or is judged to be invalid/ not infringed.
This 30 month time allows the patent holder the time to assert its patent rights in court before a
generic competitor is permitted to enter
• It was proposed by Orrin Hatch & Henry A. Waxman which became effective after 24th September 1984 which
facilitated the growth of generic industry.
• The bill altered the pharmaceutical industry substantially as it established government regulations for generic
drugs in the U.S. & made the entry of generic drugs easy.
• It established the approval pathway for generic drug products, under which applicants can submit an abbreviated
new drug application (ANDA) under section 505(j) of the Federal Food, Drug, and Cosmetic Act (FD&C Act).
• The Amendments include provisions that involve patents and exclusivities related to new drug applications, and
180-day exclusivity for certain ANDA applicants. As a general matter, the Food and Drug Administration (FDA
or the Agency) has implemented these statutory provisions within the context of application-specific decisions.
• In certain circumstances, FDA has received requests from applicants and other stakeholders for
FDA’s communications related to such decisions.
• This act brought amendments in the Patent Laws & the Federal Food, Drug & Cosmetic Act.
• The bill altered the pharma industry substantially as it established govt. regulations for generic drugs
in the USA & made it a lot easier for generic drugs to enter the market & facilitated the growth of
generic industry.
OBJECTIVES
• The original intent of the act was to increase generic drug availability in the market & make
drugs more affordable.
• The act sought to achieve the objective of increasing competition with the introduction of a
greater no. of generics in a timely manner & reward innovators with increased patent terms
during which they can enjoy market exclusivity.
• The HWA amended the FD&C act & created Abbreviated New Drug Process to submit
generic drugs to the FDA for approval.
• It also compensates the branded drug manufacturers for the time lost from the patent term
because of the regulatory approval formalities.
DRUG APPROVAL
NEW DRUG APPROVAL
GENERIC DRUG APPROVAL
• Generic Drugs are required to prove that their drug is therapeutically equivalent to innovators’
drug which they can accomplish by demonstrating bioequivalence, Pharmaceutical equivalence &
compliance with cGMP requirements.
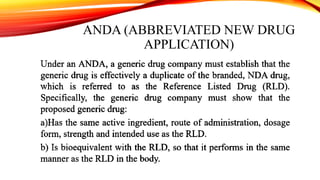
ANDA (ABBREVIATED NEW DRUG
APPLICATION)
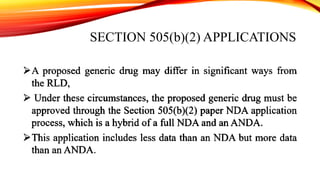
SECTION 505(b)(2) APPLICATIONS
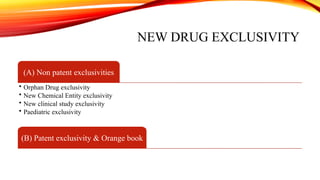
NEW DRUG EXCLUSIVITY
(A) Non patent exclusivities
• Orphan Drug exclusivity
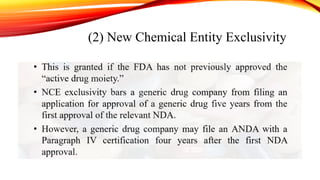
• New Chemical Entity exclusivity
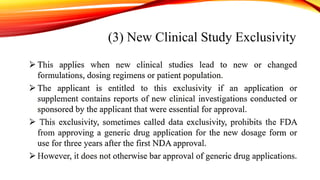
• New clinical study exclusivity
• Paediatric exclusivity
(B) Patent exclusivity & Orange book
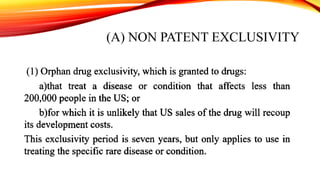
(A) NON PATENT EXCLUSIVITY
(2) New Chemical Entity Exclusivity
(3) New Clinical Study Exclusivity
(4) Paediatric Exclusivity
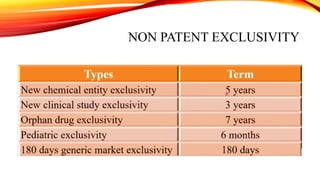
NON PATENT EXCLUSIVITY
(B) PATENT EXCLUSIVITY & ORANGE BOOK
ORANGE BOOK
• FDA publishes patent information on approved drug product in Orange product
• An NDA applicant must submit the following information about each patent:-
1. Patent no. & date on which the patent will expire
2. Type of patent i.e., drug, drug product or method of use
3. Name of patent holder
4. The name of an agent of the patent holder/ applicant
• Branded drugs listed for generics to compare with their proposed products.
CHALLENGING PATENT EXCLUSIVITY
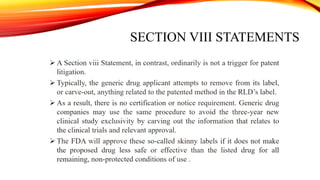
PARAGRAPH- IV CERTIFICATION
PARA IV PATENT CHALLENGE
SECTION VIII STATEMENTS
ANDA EXCLUSIVITY
PATENT TERM EXTENSION
PATENT LITIGATION UNDER THE HWAACT
BENEFITS FOR BRANDED MANUFACTURERS
• Orange book provides public notice of patents drugs
• Allows for resolution of patent disputes prior to generic entry
• 30 months stay of FDA approval of generic
• Patent term restoration
• Allows for several market exclusivities
1) Data exclusivity- 5 years for NCEs, 3 years for New clinical study exclusivity
2) Orphan drug- 7 years
3) Paediatric drugs- 6 months
LOOPHOLES OF THE ACT
AUTHORIZED GENERICS
• They are pharmaceutical products which
are approved as brand name drugs but
marketed as generics.
• Authorized generics do not bear the
trademark of the manufacturer but are
manufactured as per the brand’s
specification
WAREHOUSING PATENTS
• An innovator may patent multiple attributes
of a product (colour coating, manufacturing
process) essentially forcing the generic to
hose between waiting for the patents to
expire or file via a Paragraph IV litigation
& associated delays & cost
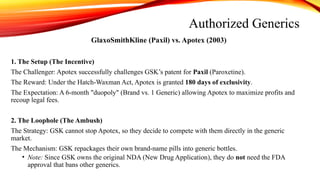
Authorized Generics
GlaxoSmithKline (Paxil) vs. Apotex (2003)
1. The Setup (The Incentive)
The Challenger: Apotex successfully challenges GSK’s patent for Paxil (Paroxetine).
The Reward: Under the Hatch-Waxman Act, Apotex is granted 180 days of exclusivity.
The Expectation: A 6-month "duopoly" (Brand vs. 1 Generic) allowing Apotex to maximize profits and
recoup legal fees.
2. The Loophole (The Ambush)
The Strategy: GSK cannot stop Apotex, so they decide to compete with them directly in the generic
market.
The Mechanism: GSK repackages their own brand-name pills into generic bottles.
• Note: Since GSK owns the original NDA (New Drug Application), they do not need the FDA
approval that bans other generics.
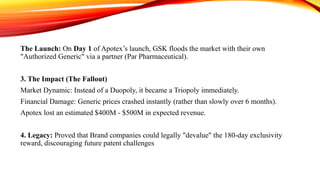
The Launch: On Day 1 of Apotex’s launch, GSK floods the market with their own
"Authorized Generic" via a partner (Par Pharmaceutical).
3. The Impact (The Fallout)
Market Dynamic: Instead of a Duopoly, it became a Triopoly immediately.
Financial Damage: Generic prices crashed instantly (rather than slowly over 6 months).
Apotex lost an estimated $400M - $500M in expected revenue.
4. Legacy: Proved that Brand companies could legally "devalue" the 180-day exclusivity
reward, discouraging future patent challenges
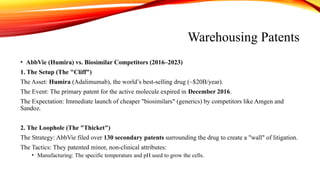
Warehousing Patents
• AbbVie (Humira) vs. Biosimilar Competitors (2016–2023)
1. The Setup (The "Cliff")
The Asset: Humira (Adalimumab), the world’s best-selling drug (~$20B/year).
The Event: The primary patent for the active molecule expired in December 2016.
The Expectation: Immediate launch of cheaper "biosimilars" (generics) by competitors like Amgen and
Sandoz.
2. The Loophole (The "Thicket")
The Strategy: AbbVie filed over 130 secondary patents surrounding the drug to create a "wall" of litigation.
The Tactics: They patented minor, non-clinical attributes:
• Manufacturing: The specific temperature and pH used to grow the cells.
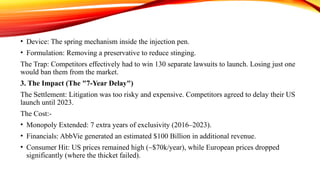
• Device: The spring mechanism inside the injection pen.
• Formulation: Removing a preservative to reduce stinging.
The Trap: Competitors effectively had to win 130 separate lawsuits to launch. Losing just one
would ban them from the market.
3. The Impact (The "7-Year Delay")
The Settlement: Litigation was too risky and expensive. Competitors agreed to delay their US
launch until 2023.
The Cost:-
• Monopoly Extended: 7 extra years of exclusivity (2016–2023).
• Financials: AbbVie generated an estimated $100 Billion in additional revenue.
• Consumer Hit: US prices remained high (~$70k/year), while European prices dropped
significantly (where the thicket failed).
CONCLUSION
• The HWA provides an expedited USFDA program for speedy generic entry & market
exclusivity.
• The HWA allows for a patent term extension of a max. of 5 years for the branded drug
manufacturer to compensate for the time lost during the NDA approval by the USFDA.