DMF of Japan and Drug
Regulatory Approval
Procedure
Presented By:
Tanisha Jain
2308212170005
Guided By:
Mrs. Zanza Patel
Assistant Professor
TABLE OF CONTENT
Application Process for Drug Product
Introduction
Timelines and Fees
General Requirements for DMF in Japan
Drug Regulatory Approval Procedure
References
The DMF System Summary
General Information about DMF of Japan
2
TABLE OF CONTENT
DMF of Japan
Timelines and Fees
Drug Regulatory Approval Procedure
References
The DMF System Summary
Items to be registered in MF
3
• Drug Master File is information registered by the
manufacturer of the API with manufacturing methods
and their drug substance to the PMDA.
• DMF is not a patent and cannot be
approved/disapproved but can be registered for
regulatory review.
• The registered data can be used as a reference in an
approval review of the drug in which the drug
substance is applied.
• It is also known as “Master File” in Japan.
INTRODUCTION:
4
General Requirements for DMF
in Japan:
Applicability of DMF in Japan:
• In Japan DMF can be submitted only
for:
• Drug Substances or intermediates (For
medical use)
• Cellular and tissue-based materials
used for manufacturing
• New and pre-mix excipients with
different composition ratios from the
existing ones.
5
General Requirements for DMF
in Japan:
Qualification of MF Holder:
• The MF holder can only be the actual
manufacturer of the API
• DMF application cannot be accepted if
the company performs only
repackaging, packaging, labelling,
storage or testing of the API.
• Customer and Contract Manufacturer
combination status.
6
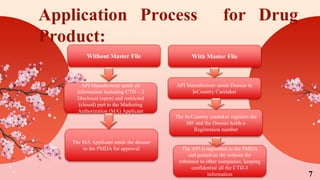
Application Process for Drug
Product:
7
Without Master File
API Manufacturer sends all
information including CTD – 3
Disclosed (open) and restricted
(closed) part to the Marketing
Authorization (MA) Applicant
The MA Applicant sends the dossier
to the PMDA for approval
The In-Country caretaker registers the
MF and the Dossier holds a
Registration number
The API is registered to the PMDA
and posted on the website for
reference to other companies, keeping
confidential all the CTD-3
information
With Master File
API Manufacturer sends Dossier to
InCountry Caretaker
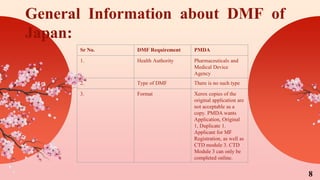
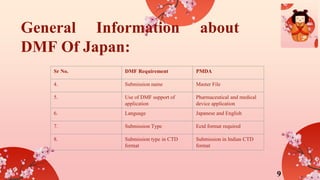
General Information about DMF of
Japan:
8
Sr No. DMF Requirement PMDA
1. Health Authority Pharmaceuticals and
Medical Device
Agency
2. Type of DMF There is no such type
3. Format Xerox copies of the
original application are
not acceptable as a
copy. PMDA wants
Application, Original
1, Duplicate 1.
Applicant for MF
Registration, as well as
CTD module 3. CTD
Module 3 can only be
completed online.
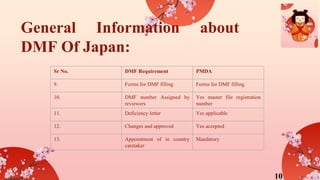
General Information about
DMF Of Japan:
9
Sr No. DMF Requirement PMDA
4. Submission name Master File
5. Use of DMF support of
application
Pharmaceutical and medical
device application
6. Language Japanese and English
7. Submission Type Ectd format required
8. Submission type in CTD
format
Submission in Indian CTD
format
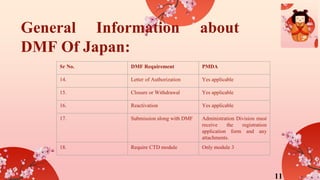
General Information about
DMF Of Japan:
10
Sr No. DMF Requirement PMDA
9. Forms for DMF filling Forms for DMF filling
10. DMF number Assigned by
reviewers
Yes master file registration
number
11. Deficiency letter Yes applicable
12. Changes and approved Yes accepted
13. Appointment of in country
caretaker
Mandatory
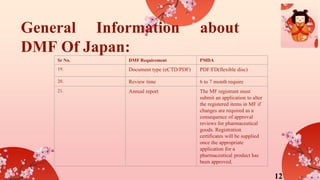
General Information about
DMF Of Japan:
Sr No. DMF Requirement PMDA
14. Letter of Authorization Yes applicable
15. Closure or Withdrawal Yes applicable
16. Reactivation Yes applicable
17. Submission along with DMF Administration Division must
receive the registration
application form and any
attachments.
18. Require CTD module Only module 3
11
General Information about
DMF Of Japan:
Sr No. DMF Requirement PMDA
19. Document type (eCTD/PDF) PDF/FD(flexible disc)
20. Review time 6 to 7 month require
21. Annual report The MF registrant must
submit an application to alter
the registered items in MF if
changes are required as a
consequence of approval
reviews for pharmaceutical
goods. Registration
certificates will be supplied
once the appropriate
application for a
pharmaceutical product has
been approved.
12
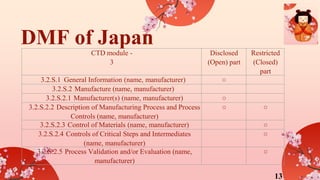
DMF of Japan
CTD module -
3
Disclosed
(Open) part
Restricted
(Closed)
part
3.2.S.1 General Information (name, manufacturer) ○
3.2.S.2 Manufacture (name, manufacturer)
3.2.S.2.1 Manufacturer(s) (name, manufacturer) ○
3.2.S.2.2 Description of Manufacturing Process and Process
Controls (name, manufacturer)
○ ○
3.2.S.2.3 Control of Materials (name, manufacturer) ○
3.2.S.2.4 Controls of Critical Steps and Intermediates
(name, manufacturer)
○
3.2.S.2.5 Process Validation and/or Evaluation (name,
manufacturer)
○
13
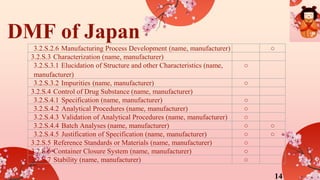
DMF of Japan
3.2.S.2.6 Manufacturing Process Development (name, manufacturer) ○
3.2.S.3 Characterization (name, manufacturer)
3.2.S.3.1 Elucidation of Structure and other Characteristics (name,
manufacturer)
○
3.2.S.3.2 Impurities (name, manufacturer) ○
3.2.S.4 Control of Drug Substance (name, manufacturer)
3.2.S.4.1 Specification (name, manufacturer) ○
3.2.S.4.2 Analytical Procedures (name, manufacturer) ○
3.2.S.4.3 Validation of Analytical Procedures (name, manufacturer) ○
3.2.S.4.4 Batch Analyses (name, manufacturer) ○ ○
3.2.S.4.5 Justification of Specification (name, manufacturer) ○ ○
3.2.S.5 Reference Standards or Materials (name, manufacturer) ○
3.2.S.6 Container Closure System (name, manufacturer) ○
3.2.S.7 Stability (name, manufacturer) ○
14
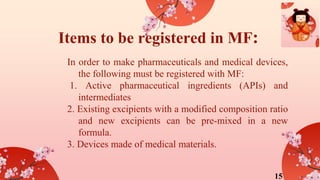
Items to be registered in MF:
In order to make pharmaceuticals and medical devices,
the following must be registered with MF:
1. Active pharmaceutical ingredients (APIs) and
intermediates
2. Existing excipients with a modified composition ratio
and new excipients can be pre-mixed in a new
formula.
3. Devices made of medical materials.
15
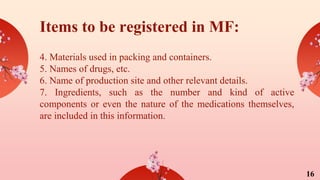
Items to be registered in MF:
4. Materials used in packing and containers.
5. Names of drugs, etc.
6. Name of production site and other relevant details.
7. Ingredients, such as the number and kind of active
components or even the nature of the medications themselves,
are included in this information.
16
Items to be registered in MF:
8. Quality control testing, manufacturing procedure, and process
control are all included in this section.
9. Specifications and test methods.
10. An expiration date and tests for stability are included.
11. A study that isn't aimed towards treating patients (mainly for
new excipients).
12. Insights on safety problems.
17
Items to be registered in MF:
13. Manufacturing operations or certification of foreign
manufacturers fall into this category.
14. The license number for a manufacturing operation, or the
date and time of the foreign manufacturer's accreditation.
15. Information about the applicant's caretaker if the manufacture
is located outside of Japan and applying for an MF registration
there.
18
● It is important for MAA/MAH, the MF Holder and the
country caretaker to understand the Japanese regulation
(PFSB/ELD Notification No. 0210001) and the guidance.
● There must be a thorough understanding of Japanese law and
instructions for the MAA/MAH, the MF Holder, and any
carers in Japan.
The DMF System Summary:
19
● The portion that has been made public. MF owner,
MAA/MAH, and the person in charge of the host nation must
all be in contact.
● Restricted (closed) area of the website The holder of the MF
and the person in charge in the host country must stay in
contact.
The DMF System Summary:
20
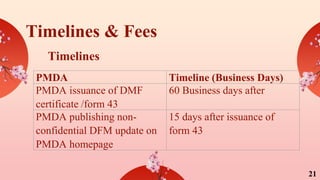
Timelines & Fees
PMDA Timeline (Business Days)
PMDA issuance of DMF
certificate /form 43
60 Business days after
PMDA publishing non-
confidential DFM update on
PMDA homepage
15 days after issuance of
form 43
Timelines
21
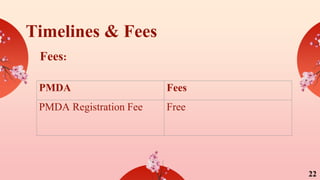
Timelines & Fees
PMDA Fees
PMDA Registration Fee Free
Fees:
22
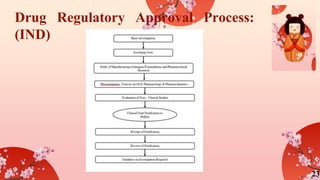
Drug Regulatory Approval Process:
(IND)
23
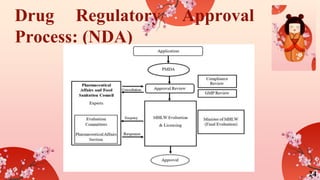
Drug Regulatory Approval
Process: (NDA)
24
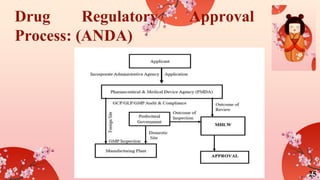
Drug Regulatory Approval
Process: (ANDA)
25
References:
1. Guidance on Drug Master File System in Japan
Master File Management Group Division of
Pharmacopoeia and Standards for Drugs Office of
Review Management.
2. Shahzeb S, Salunkhe D. STUDY ON DMF
FILLING IN CONTEXT TO US, EUROPE,
JAPAN AND ITS COMPARISON.
26
ありがとう
27