Comprehensive WHO Guidelines on Pharmaceutical Technology Transfer Processes
Detailed overview of WHO guidelines for technology transfer in pharmaceuticals, covering terminology, quality risk management, process validation, documentation, and regulatory aspects for successful drug development and commercialization.
Comprehensive WHO Guidelines on Pharmaceutical Technology Transfer Processes
1.
Prepared By
Mr. SakhareR. S.
D. Pharm., B. Pharm., B. A., M. Pharm., (Ph.D.)
Assistant Professor, Latur College Of Pharmacy.
GPAT 2020, NIPER JEE 2020, GPAT 2022, NIPER PhD Entrance 2022,
SRTMU Nanded PET 2022 Qualified
2.
WHO guidelinesfor Technology Transfer (TT):
Terminology
Technology transfer protocol
Quality risk management
Transfer from R & D to production (Process, packaging and
cleaning)
Granularity of TT Process (API, excipients, finished products,
packaging materials)
Documentation, Premises and equipments, qualification and
validation, quality control, analytical method transfer,
Approved regulatory bodies and agencies
Commercialization - practical aspects and problems (case
studies),
TT agencies in India - APCTD, NRDC, TIFAC, BCIL, TBSE
/ SIDBI
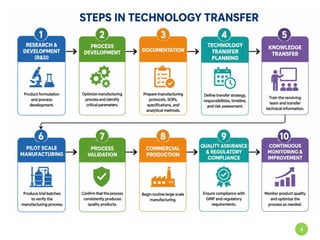
TT related documentation - confidentiality agreement,
licensing, MoUs, legal issues.
1
3.
Technology Transfer(TT):
Introduction:
Transfer of technology is defined as "a process by which a
developer of technology makes its technology available to a
commercial partner that will utilize the technology".
In pharmaceutical industry, “technology transfer” refers to the
processes of successful progress from drug discovery to
product development, clinical trials and ultimately full scale
commercialization.
Transfer of technology requires :
Documented, planned approach.
Trained and knowledgeable personnel.
Documentation of data covering all aspects of development, production and quality
control.
Usually there is a Sending unit (SU), a receiving unit (RU) and the unit managing the
process, which may or may not be a separate entity. 2
4.
Reasons for technologytransfer:
Increase Manufacturing Capacity
Lack of manufacturing capacity.
Lack of resources to launch product commercially.
Lack of marketing & distribution capability.
Exploitation in a different field of application.
Expansion into global markets.
Importance of technology transfer in pharmaceutical industry:
Ensures Successful Scale-Up
Maintains Product Quality
Reduces Manufacturing Risks
Improves Production Efficiency
Accelerates Product Commercialization
Facilitates Knowledge Transfer
Cost Reduction
Global Manufacturing Support
3
WHO GUIDELINESFOR TECHNOLOGY TRANSFER (TT)
The World Health Organization (WHO) is a specialized agency of the
United Nations looking after issues of international public health.
It was established on 7 April 1948. Its headquarter is in Geneva
Switzerland.
The WHO is a member of the United Nations development group. This
draught text was subsequently prepared by Mr. John startup United
Kingdom and by Dr. Monika Zweygarth South Africa.
The informal consultation on the World Health Organization (WHO)
guidelines for medicines quality assurance, quality control laboratories,
and transfer of technology took place from July 27–31, 2009, to review
feedback and draft subsequent revisions.
The WHO Expert Committee on Specifications for Pharmaceutical
Preparation, therefore, recommended in its 42nd report that WHO address
this issue through preparation of WHO guidelines in this area.
6
8.
TERMINOLOGY
7
Activepharmaceutical ingredient (API):
Any substance or mixture of substances intended to be used in the manufacture
of a pharmaceutical dosage form and that, when so used becomes an active
ingredient of that pharmaceutical dosage form.
Such substances are intended to furnish pharmacological activity or other
direct effect in the diagnosis, cure, mitigation, treatment, or prevention of
disease or to affect the structure and function of the body.
Good manufacturing practice:
That part of quality assurance which ensures that pharmaceutical products are
consistently produced and controlled to the quality standards appropriate to
their intended use and as required by the marketing authorization.
Process validation:
Documented evidence which provides a high degree of assurance that a
specific process will consistently result in a product that meets its
predetermined specific actions and quality characteristics.
9.
Quality Assurance(QA):
Quality assurance is a wide-ranging concept covering all matters that
individually or collectively influence the quality of a product.
It is the totality of the arrangements made with the objective of ensuring that
pharmaceutical products are of the quality required for their intended use.
Quality control(QC):
Quality control covers all measures taken, including the setting of
specifications, sampling, testing and analytical clearance, to ensure that
starting materials, intermediates, packaging materials and finished
pharmaceutical products conform with established specifications for
identity, strength, purity and other characteristics.
8
10.
Acceptance criteria:
Measurableterms under which a test result will be considered acceptable.
Change control (C/C):
A formal system by which qualified representatives of appropriate disciplines
review proposed or actual changes that might affect a validated status.
The intent is to determine the need for action that would ensure that the system
is maintained in a validated state.
Drug master file (DMF):
Detailed information concerning a specific facility, process or product
submitted to the drug regulatory authority, intended for the incorporation into
the application for marketing authorization.
Commissioning:
The setting up, adjustment and testing of equipment or a system to ensure that
it meets all the requirements, as specified in the user requirement specification,
and capacities as specified by the designer or developer.
Commissioning is carried out before qualification and validation.
9
11.
Corrective action(C/A):
Any action to be taken when the results of monitoring at a critical control
point indicate a loss of control.
Qualification:
Action of proving and documenting that any premises, systems and
equipment are properly installed, and/or work correctly and lead to the
expected results.
Qualification is often a part (the initial stage) of validation, but the
individual qualification steps alone do not constitute process validation.
Design qualification (DQ):
Documented evidence that the premises, supporting systems, utilities,
equipment and processes have been designed in accordance with the
requirements of good manufacturing practices (GMP).
10
12.
11
Installation qualification(IQ):
The performance of tests to ensure that the installations (such as machines,
measuring devices, utilities and manufacturing areas) used in a
manufacturing process are appropriately selected, correctly installed and
operate in accordance with established specifications.
Operational qualification (OQ):
Documented verification that the system or subsystem performs as intended
over all anticipated operating ranges.
Performance qualification (PQ):
Documented verification that the equipment or system operates consistently
and gives reproducibility within defined specifications and parameters for
prolonged periods.
Design space:
The multidimensional combination and interaction of input variables (e.g.
material attributes) and process parameters that have been demonstrated to
provide assurance of quality.
13.
Gap analysis:
Identification of critical elements of a process which are available at the SU
but are missing from the RU.
Good Manufacturing Practices (GMP):
That part of quality assurance which ensures that products are consistently
produced and controlled to the quality standards appropriate to their intended
use and as required by the marketing authorization.
Inter-company transfer:
A transfer of technology between sites of different companies.
Intra-company transfer:
A transfer of technology between sites of the same group of companies.
In-process control (IPC):
Checks performed during production in order to monitor and, if necessary, to
adjust the process to ensure that the product conforms to its specifications.
The control of the environment or equipment may also be regarded as a part
of in-process control.
12
14.
Quality riskmanagement (QRM):
Quality risk management is a systematic process for the assessment, control,
communication and review of risks to the quality of the pharmaceutical
product throughout the product life-cycle.
Receiving unit (RU):
The involved disciplines at an organization where a designated product,
process or method is expected to be transferred.
Sending unit (SU):
The involved disciplines at an organization from where a designated product,
process or method is expected to be transferred.
Technology transfer report (TTR):
A documented summary of a specific technology transfer project listing
procedures, acceptance criteria, results achieved and conclusions. Any
deviation should be discussed and justified.
13
15.
Validation
Itis a documented evidence that provides a high degree of assurance that a
specific process will consistently produce a product meeting its predetermined
specifications and quality attributes.
Spiking
The addition of a known amount of a compound to a standard, sample or
placebo, typically for the purpose of confirming the performance of an
analytical procedure.
Transfer of technology (TOT):
A logical procedure that controls the transfer of an established process
together with its documentation and professional expertise to site capable of
reproducing the process and its support functions to a predetermined level of
performance.
Validation master plan (VMP)
A high-level document that establishes an umbrella validation plan for the
entire project and summarizes the manufacturer's overall philosophy and
approach, to be used for establishing performance adequacy.
14
16.
Validation protocol(or plan) (VP)
A document describing the activities to be performed in a validation,
including the acceptance criteria for the approval of a manufacturing process
or a part thereof - for routine use.
Validation report (VR)
A document in which the records, results and evaluation of a completed
validation programme are assembled and summarized.
It may also contain proposals for the improvement of processes and/or
equipment
Standard operating procedure(SOP):
An authorized written procedure giving instructions for performing
operations not necessarily specific to a given product or material (e.g.
equipment operation, maintenance and cleaning, validation, cleaning of
premises and environmental control, sampling and inspection).
15
17.
Technology transfer protocol
A technology transfer protocol is a formal, written master plan that
controls and defines the systematic transfer of a manufacturing process,
analytical method, or product from a sending unit (SU) to a receiving
unit (RU).
Objectives
Ensure smooth and successful transfer of technology.
Maintain product quality, safety, and efficacy.
Demonstrate reproducibility of the manufacturing process.
Comply with regulatory and GMP requirements.
Minimize risks during scale-up and commercial production
16
18.
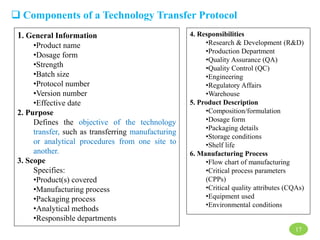
1. General Information
•Productname
•Dosage form
•Strength
•Batch size
•Protocol number
•Version number
•Effective date
2. Purpose
Defines the objective of the technology
transfer, such as transferring manufacturing
or analytical procedures from one site to
another.
3. Scope
Specifies:
•Product(s) covered
•Manufacturing process
•Packaging process
•Analytical methods
•Responsible departments
4. Responsibilities
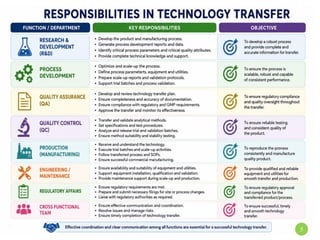
•Research & Development (R&D)
•Production Department
•Quality Assurance (QA)
•Quality Control (QC)
•Engineering
•Regulatory Affairs
•Warehouse
5. Product Description
•Composition/formulation
•Dosage form
•Packaging details
•Storage conditions
•Shelf life
6. Manufacturing Process
•Flow chart of manufacturing
•Critical process parameters
(CPPs)
•Critical quality attributes (CQAs)
•Equipment used
•Environmental conditions
Components of a Technology Transfer Protocol
17
19.
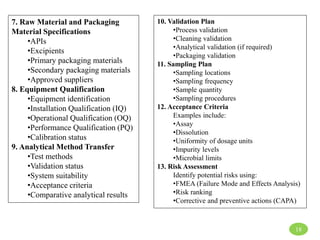
7. Raw Materialand Packaging
Material Specifications
•APIs
•Excipients
•Primary packaging materials
•Secondary packaging materials
•Approved suppliers
8. Equipment Qualification
•Equipment identification
•Installation Qualification (IQ)
•Operational Qualification (OQ)
•Performance Qualification (PQ)
•Calibration status
9. Analytical Method Transfer
•Test methods
•Validation status
•System suitability
•Acceptance criteria
•Comparative analytical results
10. Validation Plan
•Process validation
•Cleaning validation
•Analytical validation (if required)
•Packaging validation
11. Sampling Plan
•Sampling locations
•Sampling frequency
•Sample quantity
•Sampling procedures
12. Acceptance Criteria
Examples include:
•Assay
•Dissolution
•Uniformity of dosage units
•Impurity levels
•Microbial limits
13. Risk Assessment
Identify potential risks using:
•FMEA (Failure Mode and Effects Analysis)
•Risk ranking
•Corrective and preventive actions (CAPA)
18
20.
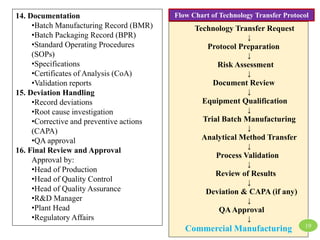
14. Documentation
•Batch ManufacturingRecord (BMR)
•Batch Packaging Record (BPR)
•Standard Operating Procedures
(SOPs)
•Specifications
•Certificates of Analysis (CoA)
•Validation reports
15. Deviation Handling
•Record deviations
•Root cause investigation
•Corrective and preventive actions
(CAPA)
•QA approval
16. Final Review and Approval
Approval by:
•Head of Production
•Head of Quality Control
•Head of Quality Assurance
•R&D Manager
•Plant Head
•Regulatory Affairs
Technology Transfer Request
↓
Protocol Preparation
↓
Risk Assessment
↓
Document Review
↓
Equipment Qualification
↓
Trial Batch Manufacturing
↓
Analytical Method Transfer
↓
Process Validation
↓
Review of Results
↓
Deviation & CAPA (if any)
↓
QAApproval
↓
Commercial Manufacturing
Flow Chart of Technology Transfer Protocol
19
21.
Quality risk management
QRM is a systematic process that is used to identify, assess. control, prevent
and mitigate all risk by appropriate and robust controls.
QRM Related guidelines: ICH Q9
Aim:
To Ensure the quality of a product
To Eliminate the risk at the patient's health.
To Ensure the Elimination of risks at the point of Origin
To protect the company from loss generated because of inadequate quality products
Principles
Risk evaluation should be based on scientific knowledge and link to the protection of the
patient
The level of Effort, formality and documentation of the QRM process should be
commensurate (adequate) with the level of risk.
20
22.
Process
1. Risk Assessment
Risk assessment is the first step where potential hazards are identified and
studied.
It consists of three sub steps.
a) Risk Identification
Recognizing potential risks that could affect product quality or patient safety.
Examples: Contamination during manufacturing, equipment failure, labeling
errors.
b) Risk Analysis
It involves both qualitative as well as quantitative analysis of the risk and
severity of risk.
Tools like FMEA (Failure Mode and Effects. Analysis), HACCP, Fault tree
Analysis may be used.
c) Risk Evaluation
It involves a comparison of Identified and analyzed risk against predefined
acceptance Criteria. 21
23.
2. Risk Control
Risk control involves deciding and implementing actions to reduce or
manage risks.
It consists of three sub-steps.
a) Risk reduction:
Introduce preventive measures.
eg: proper sterilization. Staff training, double-checking labels.
b) Risk acceptance
If risk is very low or negligible, it can be accepted.
c) Risk elimination
If risk is very critical and no control measure is effective. the activity may
be stopped.
22
24.
3. Risk Communication
Involves sharing risk information and decisions with all concerned people site
management, employees, suppliers, regulators, auditors etc.
Ensures transparency and supports good decision making.
Examples: Communicating Suppliers quality risks to manufacturing and
quality assurance teams
4. Risk Review
Risks are not static; they may Change with time, process Change or new
information
Continuous monitoring, trending and updating of risk assessments is
necessary.
Example: Reviewing risks during product lifecycle, change Control, or after
deviations.
Importance
Ensures patient Safety.
Improves product quality
Supports regulatory compliance
Provides Scientific decision-making
Optimizes resources (Efficient Resource utilization)
Supports Continuous Improvement.
Strengthens Communication and transparency: 23
25.
Tools ofQRM
1. Flowchart, Check Sheets
Simple techniques.
Used to gather and organize the data.
Facilitate decision making.
2. Fishbone | Ishikawa (Cause Effect Diagram)
Identifies possible cases of a problem.
Example: Analyzing root cause of Packaging defect.
3. Fault tree Analysis (FTA)
Top-down approach using logic diagram to find root cause of failure.
Identify System or subsystem or chain of the system failed at one time.
4. Hazard Operability Analysis (HAZOP)
This tool is based on the assumption that events are caused due to deviation in design and
operating intention.
It uses a systematic approach to identify those design and Operation intention.
5. Hazard Analysis and critical control points (HACCP)
Mainly focus on Identifying critical points from where the hazard can be originated and primary
focus is on to control hazard rather than curing it.
6. Failure Mode Effect Analysis (FMEA)
It aimed at preparing organization to detect failure during design State only by identifying all
possible failures in a design and manufacturing process. 24
26.
TT From R& D to production:
Technology transfer (TT) from R & D to production means moving the knowledge
and practices developed during research into a commercial manufacturing site in such
way that the product can be consistently produced at large scale with same quality,
safety and efficacy.
It involves following steps:
1) Process transfer:
Involves moving the complete manufacturing process (formulation, mixing,
granulation, drying, coating, etc.) developed in R & D to the production site.
Steps involved:
Preparation and review of process protocols.
Identification of critical process parameters and critical quality attributes.
Conducting pilot batches for process standardization
Scale-up of manufacturing process from laboratory to production level.
Documentation of all process steps.
25
27.
2) Packaging transfer:
Refers to replicating R&D packaging methods for commercial production while
maintaining product integrity, stability and regulatory standards.
Step involved:
Selection of suitable packaging materials.
Validation of packaging process.
Standardization of labeling and sealing process.
Assessment of packaging compatibility with the product.
3) Cleaning Transfer:
Refer cleaning procedures developed in R & D- including selection of cleaning agents and
methods are optimized and validated for production.
Development of cleaning procedures based on R&D studies.
Steps:
Validation of cleaning methods on production equipment.
Establishment of acceptance criteria for equipment cleanliness.
Documentation of cleaning processes. 26
28.
4) Documentation andcommunication
All procedures, results, and verifications should be well-documented.
Ensure coordination b/w R & D, production, quality control, and quality assurance teams
to facilitate a smooth and compliant transfer.
Example include
Technology transfer Protocol (TT protocol)
Master Batch Recorders
Packaging instructions
Cleaning validation report
Training records of staff
27