Anti Histamines Drugs
Ravish Yadav
What are histamines?
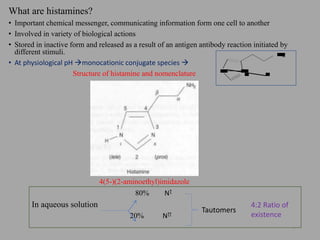
• Important chemical messenger, communicating information form one cell to another
• Involved in variety of biological actions
• Stored in inactive form and released as a result of an antigen antibody reaction initiated by
different stimuli.
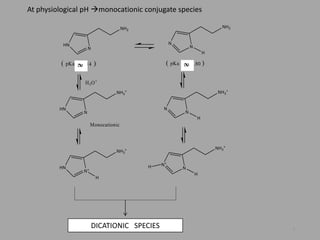
• At physiological pH monocationic conjugate species 
Structure of histamine and nomenclature
4(5-)(2-aminoethyl)imidazole
80% NƮ
In aqueous solution
20% N∏
2
4:2 Ratio of
existence
Tautomers
HN
N
NH3
+
3
HN
N
NH2
N
N
NH2
H
pKa 14( )
H3O+
pKa( )5.80
HN
N
NH3
+
N
N
NH3
+
H
Monocationic
HN
N+
NH3
+
H
N+
N
NH3
+
H
H
DICATIONIC SPECIESDICATIONIC SPECIES
At physiological pH monocationic conjugate species
Stereochemistry : Achiral, trans and gauche rotamers
4
Trans  H1 & H2 agonistic Gauche  H1 agonistic
Pharmacology of Histamine : Biosynthesis and Storage
Histidine decarboxylase
in presence of pyridoxyl phosphate
• In human : CSF, skin, bronchial and intestinal mucosa
• Storage and release : in mast cells and basophills.
Released by exocytosis as a response to immune (Ag-Ab) and non-immune
(Drug , phy. factors) stimuli.
• RECEPTORS : G - protein coupled receptors.
H1  Allergy
H2  Gastric acid secretion
H3 Neurotransmitter in CNS
5
N
HN
NH2
COOH
Histamine
N
HN
NH3
+
S-Histidine
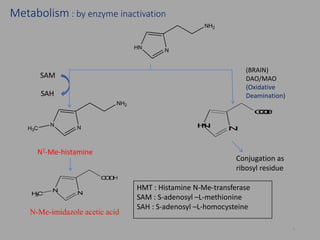
Metabolism : by enzyme inactivation
6
NHN
NH2
NN
NH2
H3C NHN
COOH
Conjugation as
ribosyl residue
NƮ-Me-histamine
SAM
SAH
(BRAIN)
DAO/MAO
(Oxidative
Deamination)
NN
COOH
H3C
N-Me-imidazole acetic acid
HMT : Histamine N-Me-transferase
SAM : S-adenosyl –L-methionine
SAH : S-adenosyl –L-homocysteine
Histamine receptors and its function: 4 different types of receptors
(G-protein coupled)
H1 : found in mammalian brain , smooth muscle from airways, GI tract,
genitourinary system, CVS, adrenal medulla & endothelial cells & lymphocytes.
• Histamine which act on H1 receptors causes inflammation
• Anti- histamines, which act on this receptor are used as anti-allergy drugs
• Molecular mass of this receptor is 56 kd & represents 487 amino acids
• Receptor contains 7 hydrophobic trans-membrane domains (TM)
• TM-3 & TM-5 are main sites for binding of H1-receptor ligands.
7
 H2 : found in gastric parietal cells, vascular smooth muscles, CNS,
Neutrophils, Heart, Uterus
• function : stimulation gastric acid secretion, regulates gastrointestinal
motility &intestinal secreation.
• Molecular mass is 40 kd and has about 359 amino acids.
• Effects of H2 receptor ligand is mediated by a stimulatory Gαs protein
coupled receptor, which in turn activates adenylate cyclase promoting the
synthesis of cAMP.
8
H3 : found in CNS, PNS, Heart, lungs, GIT, endothelial cells
• function :It is coupled to a Gi/o protein, which inhibits the action of adenylate cyclase
and regulates MAP kinase and intracellular calcium levels.
 H4 : highly expressed in bone marrow & WBC’s & regulates neutrophil
release form bone marrow
• Also expressed in small intestine, spleen, colon, liver, lungs, tonsils
• These receptor subtypes may also be involved in allergic inflammatory
responses.
9
H1 Antagonist
• MOA of H1 antagonist : Competitively inhibit the action of histamine on
tissue containing H1 receptors
• 1st Generation or classical antihistamines
• 2nd Generation antihistamines : Non-sedating, antagonistic activity at other
neurotransmitter receptors like muscarinic receptors and cardiac ion
channels.
10
1st generation : SAR:
• Diaryl substitution is essential for significant H1 receptor affinity
• Presence of two aryl rings and substituted amino moieties increases the
lipophilicity of the molecule than the endogenous agonist, Histamine.
• 2 aryl groups must be non-coplanar (not be in same plane) for effective receptor
interaction
• Basic amino group is necessary for attachment of an anionic site of the H1
receptor (N may be a simple dimethyl amino group or a part of heterocyclic ring)
• Carbon chain consists of usually 2 or 3 atoms. As a result distance between diaryl
& terminal N becomes 5 to 6 which is ideal for optimum activity
• X connecting moiety may be saturated C-O or simply C atom. This group along
with C chain, appears to serve as a spacer group for the key pharmacophoric
moieties
11
1st generation antihistamines are further classified into 5 types depending upon
connecting moiety and the nature of the aryl moieties:
 Aminoalkyl ethers (Ethanolamines)
 Ethylenediamines
 Piperazines(cyclizines)
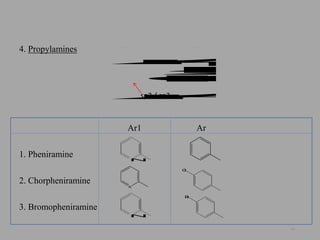
 Propylamines
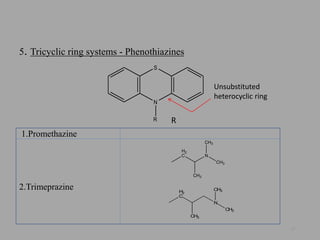
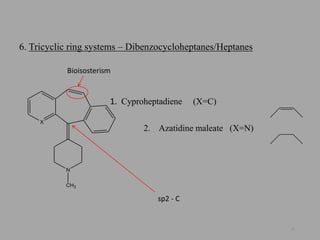
 Tricyclic ring systems (Phenothiazine and heptanes)
12
1 st generation
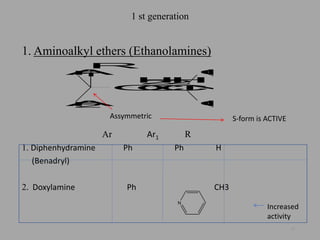
1. Aminoalkyl ethers (Ethanolamines)
Ar Ar1 R
1. Diphenhydramine Ph Ph H
(Benadryl)
2. Doxylamine Ph CH3
13
Ar1
Ar2
O
R
H2
C
H2
CN
CH3
CH3
S-form is ACTIVEAssymmetric
N
Increased
activity
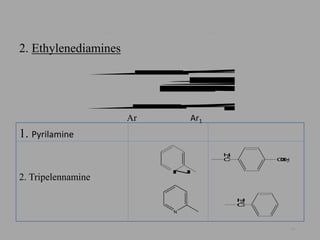
2. Ethylenediamines
Ar Ar1
1. Pyrilamine
2. Tripelennamine
14
22
3
3
r1
r
N
N
OCH3
H2
C
H2
C
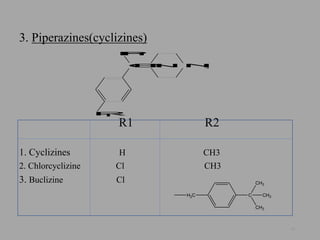
3. Piperazines(cyclizines)
R1 R2
1. Cyclizines H CH3
2. Chlorcyclizine Cl CH3
3. Buclizine Cl
15
N NR2CH
Ph
R1
H2C C
CH3
CH3
CH3
4. Propylamines
Ar1 Ar
1. Pheniramine
2. Chorpheniramine
3. Bromopheniramine
16
CH
H2
C
H2
CN
CH3
CH3
Ar
Ar1
N
N
N
Cl
Br
sp2 / sp3
5. Tricyclic ring systems - Phenothiazines
R
1.Promethazine
2.Trimeprazine
17
N
S
R
H2
C N
CH3
CH3
CH3
H2
C
N
CH3
CH3
CH3
Unsubstituted
heterocyclic ring
6. Tricyclic ring systems – Dibenzocycloheptanes/Heptanes
1. Cyproheptadiene (X=C)
2. Azatidine maleate (X=N)
18
X
N
CH3
Bioisosterism
sp2 - C
2nd Generation
1) Fexofenidine
4-[1-Hydroxy-4-[4-(hydroxyldiphenylmethyl)-1-piperinyl] butyl-α,α-
dimethyl benzeneacetic acid
19
 Fexofenidine is a primary oxidative metabolite of terfenadine
 Terfenadine is selective, long acting (>12hr) H1 antagonist with little affinity
for muscarinic, serotonergic or adrenergic receptors
 The histamine receptor affinity of this compound is believed to be related
primarily to the presence of diphenylmethyl piperidine moiety
 Terfenadine undergoes significant 1st pass metabolism, with the predominant
metabolite being fexofenidine, an active metabolite resulting from methyl
group oxidation
• When drug that inhibit this transformation such as imidazole antifungals or
macrolides, are used concurrently ,terfenadine level may rise to toxic level,
resulting in potential fatal heart rhythm problems
• Fexofenidine, like terfenadine is a selective peripheral H1 receptor ligand that
produces no clinically significant anticholinergic effect at therapeutic doses
• Fexofenadine is rapidly absorbed after oral administration
• Fexofenidine is 60-70% plasma protein bound & elimination half life is about
14 hrs
20
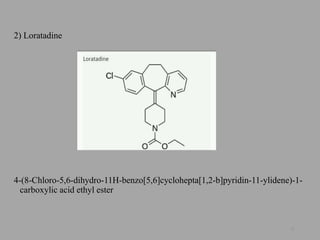
2) Loratadine
4-(8-Chloro-5,6-dihydro-11H-benzo[5,6]cyclohepta[1,2-b]pyridin-11-ylidene)-1-
carboxylic acid ethyl ester
21
• Related to tricyclic antidepressants, antihistamines azatadine and cyproheptadine
& is non sedating & neither it or nor its major metabolite, desloratidine, is
associated with potential cardiotoxic effects as that of terfenadine & astemizole
• Desloratidine is more potent H1 antagonist & more potent inhibitor of histamine
release
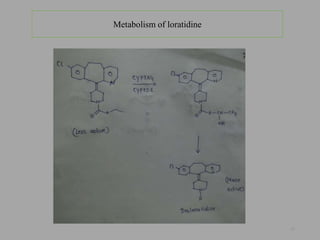
• The metabolic conversion of loratidine to descarboethoxyloratidine occurs via a
oxidative process & not via direct hydrolysis
• Both CYP2D6 & CYP3A4 appear to catalyzing this oxidative metabolic process
• Metabolite is excreted renally as a conjugate & elimination half life is about 8 to
15 hrs
22
Metabolism of loratidine
23
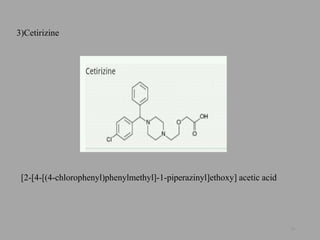
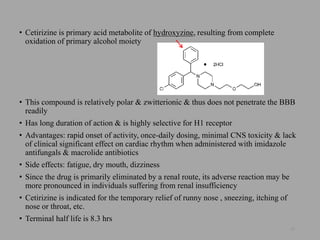
3)Cetirizine
[2-[4-[(4-chlorophenyl)phenylmethyl]-1-piperazinyl]ethoxy] acetic acid
24
• Cetirizine is primary acid metabolite of hydroxyzine, resulting from complete
oxidation of primary alcohol moiety
• This compound is relatively polar & zwitterionic & thus does not penetrate the BBB
readily
• Has long duration of action & is highly selective for H1 receptor
• Advantages: rapid onset of activity, once-daily dosing, minimal CNS toxicity & lack
of clinical significant effect on cardiac rhythm when administered with imidazole
antifungals & macrolide antibiotics
• Side effects: fatigue, dry mouth, dizziness
• Since the drug is primarily eliminated by a renal route, its adverse reaction may be
more pronounced in individuals suffering from renal insufficiency
• Cetirizine is indicated for the temporary relief of runny nose , sneezing, itching of
nose or throat, etc.
• Terminal half life is 8.3 hrs
25
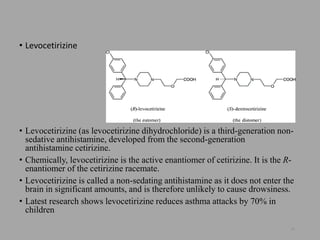
• Levocetirizine
• Levocetirizine (as levocetirizine dihydrochloride) is a third-generation non-
sedative antihistamine, developed from the second-generation
antihistamine cetirizine.
• Chemically, levocetirizine is the active enantiomer of cetirizine. It is the R-
enantiomer of the cetirizine racemate.
• Levocetirizine is called a non-sedating antihistamine as it does not enter the
brain in significant amounts, and is therefore unlikely to cause drowsiness.
• Latest research shows levocetirizine reduces asthma attacks by 70% in
children
26
4)
(E,E)-3-[6-[1-(4-methylphenyl)-3- (1-pyrrolidinyl)
-1-propenyl-2-pyridinyl]-2-propenoic acid
• Acrivastine is an analogue of triprolidine containing a carboxyethenyl moiety at the 6 position of pyridyl ring
• Acrivastine shows antihistaminic potency and duration of action comparable to those of tripolidine
• Enhanced polarity of this compound resulting from carboxyethenyl substitution limits BBB penetration & thus producing
less sedation than tripolidine
• Half life is 1.7 hr of orally administered drug
27
Triprolidine
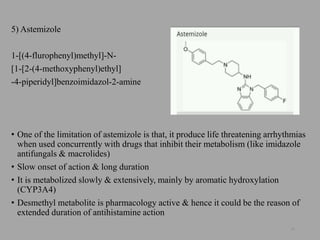
5) Astemizole
1-[(4-flurophenyl)methyl]-N-
[1-[2-(4-methoxyphenyl)ethyl]
-4-piperidyl]benzoimidazol-2-amine
• One of the limitation of astemizole is that, it produce life threatening arrhythmias
when used concurrently with drugs that inhibit their metabolism (like imidazole
antifungals & macrolides)
• Slow onset of action & long duration
• It is metabolized slowly & extensively, mainly by aromatic hydroxylation
(CYP3A4)
• Desmethyl metabolite is pharmacology active & hence it could be the reason of
extended duration of antihistamine action
28
Metabolism of astemizole
Desmethyl
(active)
HO
OH
Aromatic
hydroxylation
29
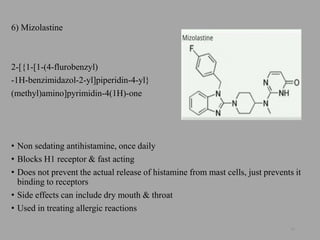
6) Mizolastine
2-[{1-[1-(4-flurobenzyl)
-1H-benzimidazol-2-yl]piperidin-4-yl}
(methyl)amino]pyrimidin-4(1H)-one
• Non sedating antihistamine, once daily
• Blocks H1 receptor & fast acting
• Does not prevent the actual release of histamine from mast cells, just prevents it
binding to receptors
• Side effects can include dry mouth & throat
• Used in treating allergic reactions
30
H2 Antagonist
31
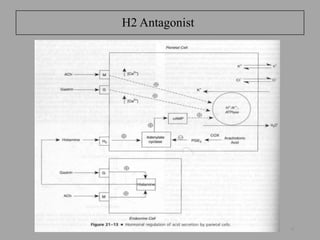
• Histamine act on H2 receptor (present on parietal cell of stomach)
which in turn activates H+/K+ ATPase system
thus more secretion of acid (H3O+) in exchange for the uptake of K+
• Anti-histamine competitively inhibit action of Histamine on H2 receptors
& prevents exchange of acid (H3O+) for K+ ,thus preventing activation of
H+/K+ ATPase system
32
Structural requirements
• H2 anti-histamines specifically designed to decrease the secretion of gastric acid
• Cimetidine, in which imidazole ring is maintained (As that of histamine). The
imidazole ring is substituted with C-4 methyl group for H2 selectivity, a 4 C side
chain includes a S atom (sulfur atom increases potency compared to C & O
congeners) & a terminal polar non-basic unit , in this case an N-cyanoguanidine
substitution (guanidine substitution with electron withdrawing groups have
significantly decreased basicity compared to guanidine and they are neutral at
physiological pH)
Histamine Cimetidine
33
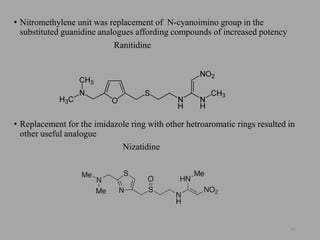
• Nitromethylene unit was replacement of N-cyanoimino group in the
substituted guanidine analogues affording compounds of increased potency
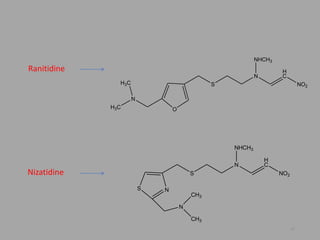
Ranitidine
• Replacement for the imidazole ring with other hetroaromatic rings resulted in
other useful analogue
Nizatidine
34
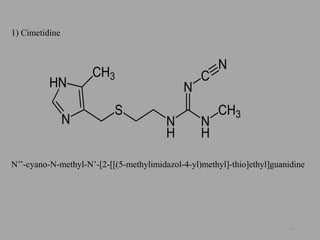
1) Cimetidine
N’’-cyano-N-methyl-N’-[2-[[(5-methylimidazol-4-yl)methyl]-thio]ethyl]guanidine
35
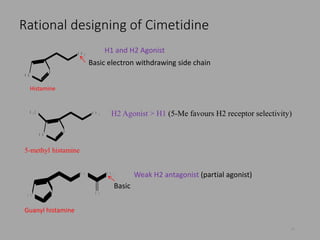
Rational designing of Cimetidine
36
N
H N
N H 2
N
H N
N H 2
H 3 C
N
H N
N H N H 2
N H
Basic electron withdrawing side chain
H1 and H2 Agonist
Histamine
5-methyl histamine
Guanyl histamine
H2 Agonist > H1 (5-Me favours H2 receptor selectivity)
Basic
Weak H2 antagonist (partial agonist)
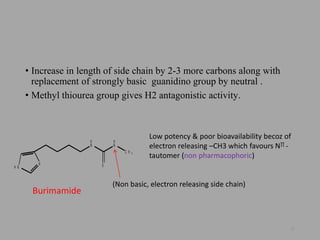
• Increase in length of side chain by 2-3 more carbons along with
replacement of strongly basic guanidino group by neutral .
• Methyl thiourea group gives H2 antagonistic activity.
37
Low potency & poor bioavailability becoz of
electron releasing –CH3 which favours N∏ -
tautomer (non pharmacophoric)
(Non basic, electron releasing side chain)
N
H N
H
N
H
N
S
C H 3
Burimamide
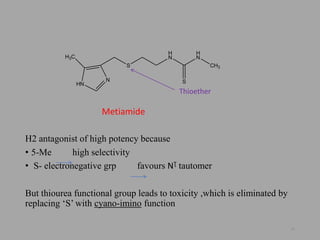
H2 antagonist of high potency because
• 5-Me high selectivity
• S- electronegative grp favours NƮ tautomer
But thiourea functional group leads to toxicity ,which is eliminated by
replacing ‘S’ with cyano-imino function
38
N
HN
S
H
N
H
N
S
CH3
H3C
Metiamide
Thioether
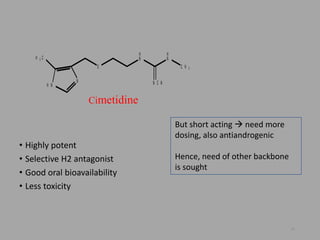
• Highly potent
• Selective H2 antagonist
• Good oral bioavailability
• Less toxicity
39
N
H N
S
H
N
H
N
N C N
C H 3
H 3 C
Cimetidine
But short acting  need more
dosing, also antiandrogenic
Hence, need of other backbone
is sought
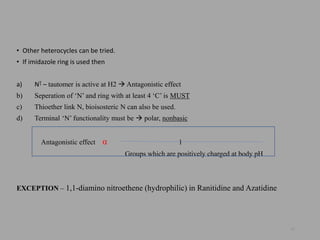
• Other heterocycles can be tried.
• If imidazole ring is used then
a) NƮ – tautomer is active at H2  Antagonistic effect
b) Seperation of ‘N’ and ring with at least 4 ‘C’ is MUST
c) Thioether link N, bioisosteric N can also be used.
d) Terminal ‘N’ functionality must be  polar, nonbasic
Antagonistic effect α 1
Groups which are positively charged at body pH
EXCEPTION – 1,1-diamino nitroethene (hydrophilic) in Ranitidine and Azatidine
40
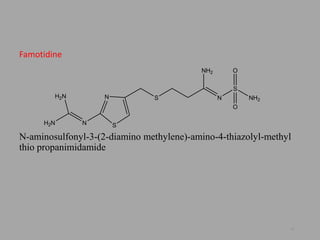
Famotidine
N-aminosulfonyl-3-(2-diamino methylene)-amino-4-thiazolyl-methyl
thio propanimidamide
41
N
S
S N
S
NH2
NH2N
H2N
NH2 O
O
O
S
N
H
C
NO2
N
H3C
H3C
NHCH3
42
NS
S
N
H
C
NO2
NHCH3
N
CH3
CH3
Nizatidine
Ranitidine
• Cimetidine reduces hepatic metabolism of drugs biotransformed by CYP450,
delaying elimination & increasing serum levels of these drugs
• Concominant therapy of patients with cimetidine & drugs metabolized by
hepatic microsomal enzymes, particularly those of low therapeutic ratio or in
patients with renal or hepatic impairment, may require dosage adjustment
• If concurrent azole therapy is required, it is best to administer it at least 2 hrs
before Cimetidine administration.
• Has weak antiandrogenic effect
• High oral bioavailability with plasma half life of about 2 hrs which is increased
in renal or hepatic impairment & in elderly
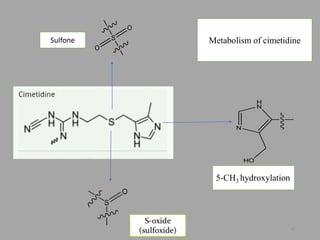
• Cimetidine is metabolized (S-oxidation, 5-CH3hydroxylation) & eliminated by
renal excretion
43
Metabolism of cimetidineS
O
O
Sulfone
N
H
N
HO
5-CH3 hydroxylation
44
2) Ranitidine
N-[2-[[[5-(dimethylamino)methyl]-2-furanyl]methyl]thio]ethyl]-N’-methyl-2-
nitro-1,1-ethenediamine
• Bioavailability of an oral dose is 50 -60% & is not affected by presence of food
• Some antacid may reduce it’s absorption & should not be taken within 1hr
administration of the H2 blocker
• Plasma half life is about 2-3hrs & metabolites are excreted in urine
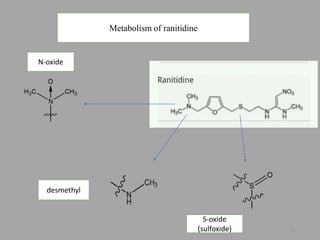
• 3 metabolites, ranitidine N-oxide, ranitidine S-oxide & desmethyl ranitidine have
been identified
• Weak inhibitor of hepatic CYP450 mixed function oxidase system
• Ranitidine is used (as bismuth citrate) with macrolide antibiotics
(clarithromycin) in treating patients with an active duodenal ulcer associated
with H.pylori infection
45
Metabolism of ranitidine
N-oxide
desmethyl
S-oxide
(sulfoxide) 46
3) Famotidine
N’-(aminosulfonyl)-3-[[[2-(diaminomethylene)-amino]-4-thiazolyl]methyl]thio]
propanimidamide
• Famotidine is a competitive inhibitor of H2 receptors & inhibits basal &
nocturnal gastric secretion as well as secretion stimulated by food & pentagastrin
• Used for short term treatment of duodenal & benign gastric ulcers, GERD,
pathological hypersecretory conditions ( eg. Zollinger-Ellison syndrome) &
heartburn
• Studies with Famotidine in humans, in animals models & in vitro have shown no
significant interference with the disposition of compounds metabolized by the
hepatic microsomal enzymes (eg. CYP450 system)
47
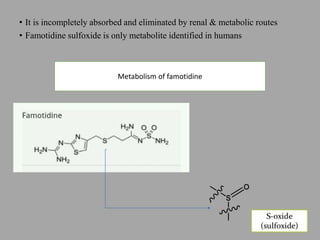
• It is incompletely absorbed and eliminated by renal & metabolic routes
• Famotidine sulfoxide is only metabolite identified in humans
48
Metabolism of famotidine
4) Nizatidine
N-[2-[[[2-(dimethylamino)-methyl]-4-thiazolyl]methyl]thio] ethyl]-N’-methyl
-2-nitro-1,1-ethenediamine
• Nizatidine has excellent oral bioavailability (>90%) & effects of antacids &
food on its bioavailability are not clinically significant
• Elimination half life is 1-2 hrs
• Excreted primarily in urine & mostly as unchanged drug
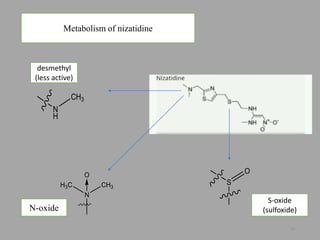
• Metabolites include nizatidine sulfoxide, N-desmethylnizatidine & nizatidine N-
oxide
• No antiandrogenic action or inhibitory effects on CYP450-linked drug
metabolizing enzyme system
49
Metabolism of nizatidine
desmethyl
(less active)
N
CH3H3C
O
N-oxide
N
H
CH3
S
O
S-oxide
(sulfoxide)
50
Proton Pump Inhibitor (PPI)
• The final step in acid secretion in parietal cells of the gastric mucosa is a
process mediated by H+/K+ATPase, the gastric proton pump which catalyzes
the exchange of hydrogen ions for potassium ions
• PPI inhibits gastric acid secretion irrespective of receptor stimulation process
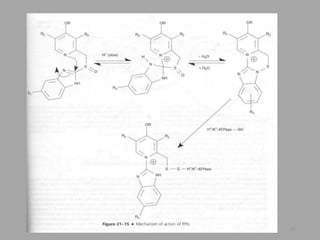
• These agents have irreversible effects on the secretion of gastric acid, because
molecule rearrange in strongly acidic environment of parietal cell
• Covalent bonding of rearranged inhibitor to H+/K+ATPase results in
inactivation of catalytic function of proton pump
• One of the site is cystein-813 & these cysteins are in different environment &
different PPI’s bind differentially to them & other sulfhydryl groups
• In covalent binding, disulfide bonds are formed with receptor
51
52
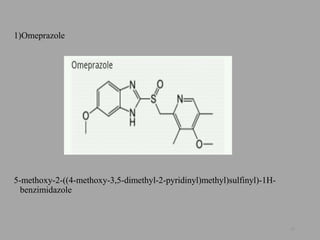
1)Omeprazole
5-methoxy-2-((4-methoxy-3,5-dimethyl-2-pyridinyl)methyl)sulfinyl)-1H-
benzimidazole
53
• Omeprazole is an amphoteric compound & is acid labile, hence, it is
formulated as delayed release capsule containing enteric coated granule
• Plasma half life is about 1 hr
• Most of an oral dose of omeprazole is excreted in the urine as metabolites with
insignificant antisecretory activity
• The primary metabolites of omeprazole are 5-hydroxyomeprazole (by
CYP2C19) and omeprazole sulfone (CYP3A4)
• The antisecretory actions of omeprazole persists 24-72 hrs, long after the drug
has disappeared from plasma, which is consistent with its suggested mechanism
of action involving irreversible inhibition of proton pump, H+/K+ ATPase.
• Used in treatment of heartburn, duodenal ulcer, gastric ulcer, etc.
54
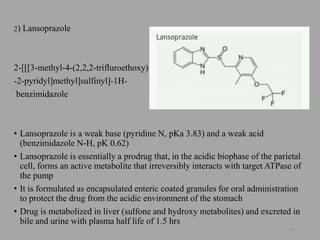
2) Lansoprazole
2-[[[3-methyl-4-(2,2,2-trifluroethoxy)
-2-pyridyl]methyl]sulfinyl]-1H-
benzimidazole
• Lansoprazole is a weak base (pyridine N, pKa 3.83) and a weak acid
(benzimidazole N-H, pK 0.62)
• Lansoprazole is essentially a prodrug that, in the acidic biophase of the parietal
cell, forms an active metabolite that irreversibly interacts with target ATPase of
the pump
• It is formulated as encapsulated enteric coated granules for oral administration
to protect the drug from the acidic environment of the stomach
• Drug is metabolized in liver (sulfone and hydroxy metabolites) and excreted in
bile and urine with plasma half life of 1.5 hrs
55
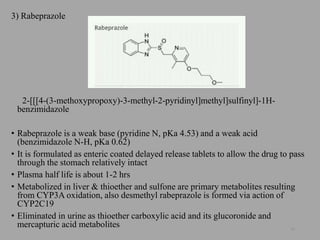
3) Rabeprazole
2-[[[4-(3-methoxypropoxy)-3-methyl-2-pyridinyl]methyl]sulfinyl]-1H-
benzimidazole
• Rabeprazole is a weak base (pyridine N, pKa 4.53) and a weak acid
(benzimidazole N-H, pKa 0.62)
• It is formulated as enteric coated delayed release tablets to allow the drug to pass
through the stomach relatively intact
• Plasma half life is about 1-2 hrs
• Metabolized in liver & thioether and sulfone are primary metabolites resulting
from CYP3A oxidation, also desmethyl rabeprazole is formed via action of
CYP2C19
• Eliminated in urine as thioether carboxylic acid and its glucoronide and
mercapturic acid metabolites 56
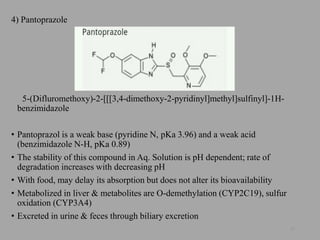
4) Pantoprazole
5-(Difluromethoxy)-2-[[[3,4-dimethoxy-2-pyridinyl]methyl]sulfinyl]-1H-
benzimidazole
• Pantoprazol is a weak base (pyridine N, pKa 3.96) and a weak acid
(benzimidazole N-H, pKa 0.89)
• The stability of this compound in Aq. Solution is pH dependent; rate of
degradation increases with decreasing pH
• With food, may delay its absorption but does not alter its bioavailability
• Metabolized in liver & metabolites are O-demethylation (CYP2C19), sulfur
oxidation (CYP3A4)
• Excreted in urine & feces through biliary excretion
57
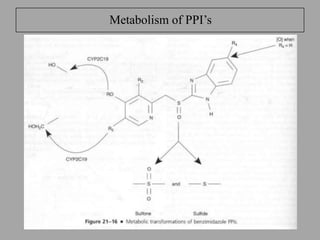
Metabolism of PPI’s
58
59
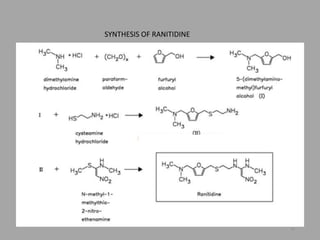
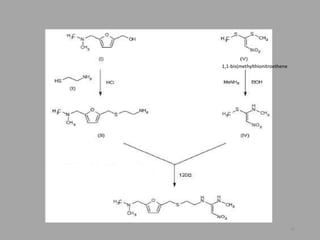
SYNTHESIS OF RANITIDINE
60
1,1-bis(methylthionitroethene
References:
•Foye’s Principles of Medicinal Chemistry.
•Wilson & Grisvold’s Textbook of Organic Medicinal
and Pharmaceutical Chemistry.
•Textbook of Medicinal Chemistry (vol-1); K.Ilango &
P.Valentina(For synthesis only).
61